[English] 日本語
Yorodumi
- PDB-4i3h: A three-gate structure of topoisomerase IV from Streptococcus pne... -

+
Open data


ID or keywords:

Loading...

-
Basic information

Entry
Database: PDB / ID: 4i3h
TitleA three-gate structure of topoisomerase IV from Streptococcus pneumoniae
Components
  • DNA (5'-D(*CP*AP*AP*AP*GP*GP*CP*GP*GP*TP*AP*AP*TP*AP*CP*GP*GP*TP*TP*AP*TP*CP*CP*AP*CP*AP*GP*AP*AP*TP*CP*AP*GP*G)-3')
  • DNA (5'-D(*CP*CP*TP*GP*AP*TP*TP*CP*TP*GP*TP*GP*GP*AP*TP*AP*AP*CP*CP*GP*TP*AP*TP*TP*AP*CP*CP*GP*CP*CP*TP*TP*TP*G)-3')
  • DNA topoisomerase 4 subunit B,DNA topoisomerase 4 subunit A
KeywordsISOMERASE/DNA / DNA unwinding / supercoiling / ISOMERASE-DNA complex
Function / homology
Function and homology information


DNA topoisomerase type II (double strand cut, ATP-hydrolyzing) complex / DNA negative supercoiling activity / DNA topoisomerase (ATP-hydrolysing) / extrinsic component of plasma membrane / DNA topological change / chromosome segregation / chromosome / DNA binding / ATP binding / cytoplasm
Similarity search - Function
DNA topoisomerase IV subunit A, Gram-positive / DNA topoisomerase 4 subunit B, Firmicutes/Mollicutes / Rossmann fold - #670 / DNA gyrase/topoisomerase IV, subunit A, C-terminal repeat / DNA gyrase/topoisomerase IV, subunit A, C-terminal / DNA gyrase C-terminal domain, beta-propeller / : / DNA topoisomerase, type IIA, subunit B / DNA gyrase B subunit, C-terminal / DNA gyrase B subunit, carboxyl terminus ...DNA topoisomerase IV subunit A, Gram-positive / DNA topoisomerase 4 subunit B, Firmicutes/Mollicutes / Rossmann fold - #670 / DNA gyrase/topoisomerase IV, subunit A, C-terminal repeat / DNA gyrase/topoisomerase IV, subunit A, C-terminal / DNA gyrase C-terminal domain, beta-propeller / : / DNA topoisomerase, type IIA, subunit B / DNA gyrase B subunit, C-terminal / DNA gyrase B subunit, carboxyl terminus / Topoisomerase (Topo) IIA-type catalytic domain profile. / DNA topoisomerase, type IIA, alpha-helical domain superfamily / DNA topoisomerase, type IIA, domain A / DNA topoisomerase, type IIA, domain A, alpha-beta / DNA gyrase/topoisomerase IV, subunit A / DNA Topoisomerase IV / DNA topoisomerase, type IIA, subunit B, domain 2 / DNA gyrase B / DNA topoisomerase, type IIA / DNA topoisomerase, type IIA, conserved site / DNA topoisomerase II signature. / TopoisomeraseII / DNA topoisomerase, type IIA, subunit B, C-terminal / Toprim domain / DNA topoisomerase, type IIA-like domain superfamily / Ribosomal Protein S5; domain 2 - #10 / Toprim domain profile. / TOPRIM domain / Ribosomal Protein S5; domain 2 / Histidine kinase-like ATPase, C-terminal domain / Heat Shock Protein 90 / Histidine kinase-, DNA gyrase B-, and HSP90-like ATPase / Histidine kinase-like ATPases / Histidine kinase/HSP90-like ATPase / Histidine kinase/HSP90-like ATPase superfamily / Ribosomal protein S5 domain 2-type fold, subgroup / Ribosomal protein S5 domain 2-type fold / Rossmann fold / 2-Layer Sandwich / 3-Layer(aba) Sandwich / Alpha Beta
Similarity search - Domain/homology
DNA / DNA (> 10) / DNA topoisomerase 4 subunit A / DNA topoisomerase 4 subunit B
Similarity search - Component
Biological speciesStreptococcus pneumoniae TIGR4 (bacteria)
synthetic construct (others)
MethodX-RAY DIFFRACTION / SYNCHROTRON / MOLECULAR REPLACEMENT / Resolution: 3.7 Å
AuthorsRamaswamy, V. / Laponogov, I. / Veselkov, D.A. / Pan, X.S. / Crevel, I. / Fisher, L.M. / Sanderson, M.R.
Funding support United Kingdom, 1items
OrganizationGrant numberCountry
Biotechnology and Biological Sciences Research Council (BBSRC)BB/K010069/1 United Kingdom
CitationJournal: Nucleic Acids Res. / Year: 2013
Title: Structure of an 'open' clamp type II topoisomerase-DNA complex provides a mechanism for DNA capture and transport.
Authors: Laponogov, I. / Veselkov, D.A. / Crevel, I.M. / Pan, X.S. / Fisher, L.M. / Sanderson, M.R.
History
DepositionNov 26, 2012Deposition site: RCSB / Processing site: RCSB
Revision 1.0Aug 28, 2013Provider: repository / Type: Initial release
Revision 1.1Sep 4, 2013Group: Database references
Revision 1.2Dec 4, 2013Group: Database references
Revision 1.3Jul 26, 2017Group: Data collection / Refinement description / Source and taxonomy
Category: diffrn_detector / entity_src_gen / software / Item: _diffrn_detector.detector
Revision 1.4Sep 20, 2023Group: Data collection / Database references ...Data collection / Database references / Derived calculations / Refinement description
Category: chem_comp_atom / chem_comp_bond ...chem_comp_atom / chem_comp_bond / database_2 / pdbx_initial_refinement_model / pdbx_struct_conn_angle / struct_conn / struct_ref_seq_dif / struct_site
Item: _database_2.pdbx_DOI / _database_2.pdbx_database_accession ..._database_2.pdbx_DOI / _database_2.pdbx_database_accession / _pdbx_struct_conn_angle.ptnr1_auth_asym_id / _pdbx_struct_conn_angle.ptnr1_auth_comp_id / _pdbx_struct_conn_angle.ptnr1_auth_seq_id / _pdbx_struct_conn_angle.ptnr1_label_asym_id / _pdbx_struct_conn_angle.ptnr1_label_comp_id / _pdbx_struct_conn_angle.ptnr1_label_seq_id / _pdbx_struct_conn_angle.ptnr2_auth_asym_id / _pdbx_struct_conn_angle.ptnr2_label_asym_id / _pdbx_struct_conn_angle.ptnr3_auth_asym_id / _pdbx_struct_conn_angle.ptnr3_auth_comp_id / _pdbx_struct_conn_angle.ptnr3_auth_seq_id / _pdbx_struct_conn_angle.ptnr3_label_asym_id / _pdbx_struct_conn_angle.ptnr3_label_comp_id / _pdbx_struct_conn_angle.ptnr3_label_seq_id / _pdbx_struct_conn_angle.value / _struct_conn.pdbx_dist_value / _struct_conn.ptnr1_auth_asym_id / _struct_conn.ptnr1_auth_comp_id / _struct_conn.ptnr1_auth_seq_id / _struct_conn.ptnr1_label_asym_id / _struct_conn.ptnr1_label_comp_id / _struct_conn.ptnr1_label_seq_id / _struct_conn.ptnr2_auth_asym_id / _struct_conn.ptnr2_label_asym_id / _struct_ref_seq_dif.details / _struct_site.pdbx_auth_asym_id / _struct_site.pdbx_auth_comp_id / _struct_site.pdbx_auth_seq_id
Revision 2.0Sep 2, 2026Group: Advisory / Atomic model ...Advisory / Atomic model / Author supporting evidence / Data collection / Database references / Derived calculations / Other / Polymer sequence / Refinement description / Source and taxonomy / Structure summary
Category: atom_site / atom_site_anisotrop ...atom_site / atom_site_anisotrop / atom_type / audit_author / cell / database_PDB_matrix / diffrn / entity / entity_name_com / entity_poly / entity_poly_seq / entity_src_gen / ndb_struct_conf_na / ndb_struct_na_base_pair / ndb_struct_na_base_pair_step / pdbx_audit_support / pdbx_contact_author / pdbx_database_status / pdbx_entity_src_syn / pdbx_entry_details / pdbx_nonpoly_scheme / pdbx_poly_seq_scheme / pdbx_refine_tls / pdbx_refine_tls_group / pdbx_struct_assembly / pdbx_struct_conn_angle / pdbx_struct_sheet_hbond / pdbx_unobs_or_zero_occ_atoms / pdbx_unobs_or_zero_occ_residues / pdbx_validate_close_contact / pdbx_validate_peptide_omega / pdbx_validate_rmsd_angle / pdbx_validate_rmsd_bond / pdbx_validate_torsion / refine / refine_hist / refine_ls_restr / refine_ls_shell / reflns_shell / software / space_group / space_group_symop / struct / struct_asym / struct_biol / struct_conf / struct_conn / struct_conn_type / struct_mon_prot_cis / struct_ref / struct_ref_seq / struct_ref_seq_dif / struct_sheet / struct_sheet_order / struct_sheet_range / struct_site / struct_site_gen / symmetry
Item: _cell.volume / _diffrn.pdbx_serial_crystal_experiment ..._cell.volume / _diffrn.pdbx_serial_crystal_experiment / _entity.details / _entity.formula_weight / _entity.pdbx_description / _entity.pdbx_ec / _entity.src_method / _entity_poly.pdbx_seq_one_letter_code / _entity_poly.pdbx_seq_one_letter_code_can / _entity_poly.pdbx_strand_id / _entity_poly.type / _entity_poly_seq.entity_id / _entity_poly_seq.mon_id / _entity_poly_seq.num / _entity_src_gen.entity_id / _entity_src_gen.gene_src_strain / _entity_src_gen.pdbx_end_seq_num / _entity_src_gen.pdbx_gene_src_gene / _entity_src_gen.pdbx_gene_src_scientific_name / _entity_src_gen.pdbx_seq_type / _ndb_struct_na_base_pair.buckle / _ndb_struct_na_base_pair.hbond_type_12 / _ndb_struct_na_base_pair.hbond_type_28 / _ndb_struct_na_base_pair.i_auth_asym_id / _ndb_struct_na_base_pair.i_auth_seq_id / _ndb_struct_na_base_pair.i_label_asym_id / _ndb_struct_na_base_pair.i_label_comp_id / _ndb_struct_na_base_pair.i_label_seq_id / _ndb_struct_na_base_pair.j_auth_asym_id / _ndb_struct_na_base_pair.j_auth_seq_id / _ndb_struct_na_base_pair.j_label_asym_id / _ndb_struct_na_base_pair.j_label_comp_id / _ndb_struct_na_base_pair.j_label_seq_id / _ndb_struct_na_base_pair.opening / _ndb_struct_na_base_pair.pair_name / _ndb_struct_na_base_pair.propeller / _ndb_struct_na_base_pair.shear / _ndb_struct_na_base_pair.stagger / _ndb_struct_na_base_pair.stretch / _ndb_struct_na_base_pair_step.helical_rise / _ndb_struct_na_base_pair_step.helical_twist / _ndb_struct_na_base_pair_step.i_auth_asym_id_1 / _ndb_struct_na_base_pair_step.i_auth_asym_id_2 / _ndb_struct_na_base_pair_step.i_auth_seq_id_1 / _ndb_struct_na_base_pair_step.i_auth_seq_id_2 / _ndb_struct_na_base_pair_step.i_label_asym_id_1 / _ndb_struct_na_base_pair_step.i_label_asym_id_2 / _ndb_struct_na_base_pair_step.i_label_comp_id_1 / _ndb_struct_na_base_pair_step.i_label_comp_id_2 / _ndb_struct_na_base_pair_step.i_label_seq_id_1 / _ndb_struct_na_base_pair_step.i_label_seq_id_2 / _ndb_struct_na_base_pair_step.inclination / _ndb_struct_na_base_pair_step.j_auth_asym_id_1 / _ndb_struct_na_base_pair_step.j_auth_asym_id_2 / _ndb_struct_na_base_pair_step.j_auth_seq_id_1 / _ndb_struct_na_base_pair_step.j_auth_seq_id_2 / _ndb_struct_na_base_pair_step.j_label_asym_id_1 / _ndb_struct_na_base_pair_step.j_label_asym_id_2 / _ndb_struct_na_base_pair_step.j_label_comp_id_1 / _ndb_struct_na_base_pair_step.j_label_comp_id_2 / _ndb_struct_na_base_pair_step.j_label_seq_id_1 / _ndb_struct_na_base_pair_step.j_label_seq_id_2 / _ndb_struct_na_base_pair_step.rise / _ndb_struct_na_base_pair_step.roll / _ndb_struct_na_base_pair_step.shift / _ndb_struct_na_base_pair_step.slide / _ndb_struct_na_base_pair_step.step_name / _ndb_struct_na_base_pair_step.tilt / _ndb_struct_na_base_pair_step.tip / _ndb_struct_na_base_pair_step.twist / _ndb_struct_na_base_pair_step.x_displacement / _ndb_struct_na_base_pair_step.y_displacement / _pdbx_database_status.SG_entry / _pdbx_nonpoly_scheme.auth_seq_num / _pdbx_nonpoly_scheme.pdb_strand_id / _pdbx_poly_seq_scheme.asym_id / _pdbx_poly_seq_scheme.auth_mon_id / _pdbx_poly_seq_scheme.auth_seq_num / _pdbx_poly_seq_scheme.entity_id / _pdbx_poly_seq_scheme.mon_id / _pdbx_poly_seq_scheme.ndb_seq_num / _pdbx_poly_seq_scheme.pdb_mon_id / _pdbx_poly_seq_scheme.pdb_seq_num / _pdbx_poly_seq_scheme.pdb_strand_id / _pdbx_poly_seq_scheme.seq_id / _pdbx_struct_assembly.details / _pdbx_validate_close_contact.auth_asym_id_2 / _pdbx_validate_close_contact.auth_atom_id_1 / _pdbx_validate_close_contact.auth_atom_id_2 / _pdbx_validate_close_contact.auth_comp_id_1 / _pdbx_validate_close_contact.auth_comp_id_2 / _pdbx_validate_close_contact.auth_seq_id_1 / _pdbx_validate_close_contact.auth_seq_id_2 / _pdbx_validate_close_contact.dist / _refine.B_iso_mean / _refine.aniso_B[1][1] / _refine.aniso_B[1][2] / _refine.aniso_B[1][3] / _refine.aniso_B[2][2] / _refine.aniso_B[2][3] / _refine.aniso_B[3][3] / _refine.details / _refine.ls_R_factor_R_free / _refine.ls_R_factor_R_work / _refine.ls_R_factor_obs / _refine.ls_d_res_low / _refine.ls_number_reflns_R_work / _refine.ls_number_reflns_all / _refine.ls_number_reflns_obs / _refine.overall_SU_ML / _refine.pdbx_ls_cross_valid_method / _refine.pdbx_ls_sigma_F / _refine.pdbx_overall_phase_error / _refine.pdbx_solvent_shrinkage_radii / _refine.pdbx_solvent_vdw_probe_radii / _refine.pdbx_stereochemistry_target_values / _refine.solvent_model_param_bsol / _refine.solvent_model_param_ksol / _refine_hist.d_res_low / _refine_hist.number_atoms_total / _refine_hist.pdbx_number_atoms_nucleic_acid / _refine_hist.pdbx_number_atoms_protein / _refine_ls_restr.dev_ideal / _refine_ls_restr.number / _refine_ls_restr.type / _refine_ls_shell.R_factor_R_free / _refine_ls_shell.R_factor_R_work / _refine_ls_shell.d_res_high / _refine_ls_shell.d_res_low / _refine_ls_shell.number_reflns_R_work / _refine_ls_shell.percent_reflns_obs / _reflns_shell.number_unique_obs / _software.classification / _software.name / _software.version / _struct.pdbx_CASP_flag / _struct_asym.entity_id / _struct_ref.db_code / _struct_ref.entity_id / _struct_ref.pdbx_align_begin / _struct_ref.pdbx_db_accession / _struct_ref.pdbx_seq_one_letter_code / _struct_ref_seq.db_align_beg / _struct_ref_seq.db_align_end / _struct_ref_seq.pdbx_auth_seq_align_beg / _struct_ref_seq.pdbx_auth_seq_align_end / _struct_ref_seq.pdbx_db_accession / _struct_ref_seq.pdbx_strand_id / _struct_ref_seq.ref_id / _symmetry.space_group_name_Hall
Description: Atomic clashes
Details: This structure has been re-refined to improve the model in the following ways: 1. Fix the twisted peptide and non-proline cis-peptide linkages 2. Reduce the Clashscore to acceptable range. 3. ...Details: This structure has been re-refined to improve the model in the following ways: 1. Fix the twisted peptide and non-proline cis-peptide linkages 2. Reduce the Clashscore to acceptable range. 3. Reduce the Ramachandran outliers and increase number of Ramachandran favoured residues to an acceptable range. 4. Reduce the Rotamer outliers and increase number of residues with favoured Rotamers to an acceptable range.
Provider: author / Type: Coordinate replacement

-
Structure visualization

Structure viewerMolecule:
MolmilJmol/JSmol

Downloads & links

-
Assembly

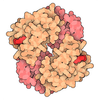
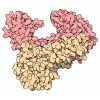
Deposited unit
A: DNA topoisomerase 4 subunit B,DNA topoisomerase 4 subunit A
B: DNA topoisomerase 4 subunit B,DNA topoisomerase 4 subunit A
G: DNA (5'-D(*CP*AP*AP*AP*GP*GP*CP*GP*GP*TP*AP*AP*TP*AP*CP*GP*GP*TP*TP*AP*TP*CP*CP*AP*CP*AP*GP*AP*AP*TP*CP*AP*GP*G)-3')
H: DNA (5'-D(*CP*CP*TP*GP*AP*TP*TP*CP*TP*GP*TP*GP*GP*AP*TP*AP*AP*CP*CP*GP*TP*AP*TP*TP*AP*CP*CP*GP*CP*CP*TP*TP*TP*G)-3')
E: DNA (5'-D(*CP*AP*AP*AP*GP*GP*CP*GP*GP*TP*AP*AP*TP*AP*CP*GP*GP*TP*TP*AP*TP*CP*CP*AP*CP*AP*GP*AP*AP*TP*CP*AP*GP*G)-3')
F: DNA (5'-D(*CP*CP*TP*GP*AP*TP*TP*CP*TP*GP*TP*GP*GP*AP*TP*AP*AP*CP*CP*GP*TP*AP*TP*TP*AP*CP*CP*GP*CP*CP*TP*TP*TP*G)-3')
hetero molecules


Theoretical massNumber of molelcules
Total (without water)298,5569
Polymers298,4836
Non-polymers733
Water362
1


  • Idetical with deposited unit
  • defined by author&software
TypeNameSymmetry operationNumber
identity operation1_555x,y,z1
Unit cell
Length a, b, c (Å)160.600, 160.600, 280.560
Angle α, β, γ (deg.)90.000, 90.000, 90.000
Int Tables number94
Space group name H-MP42212
Space group name HallP4n2n
Symmetry operation#1: x,y,z
#2: -y+1/2,x+1/2,z+1/2
#3: y+1/2,-x+1/2,z+1/2
#4: x+1/2,-y+1/2,-z+1/2
#5: -x+1/2,y+1/2,-z+1/2
#6: -x,-y,z
#7: y,x,-z
#8: -y,-x,-z

-
Components

#1: Protein DNA topoisomerase 4 subunit B,DNA topoisomerase 4 subunit A / Topoisomerase IV subunit A


Mass: 128323.859 Da / Num. of mol.: 2
Source method: isolated from a genetically manipulated source
Source: (gene. exp.) Streptococcus pneumoniae TIGR4 (bacteria)
Gene: parE, parC, SP_0855 / Production host: Escherichia coli (E. coli)
References: UniProt: Q59961, UniProt: P72525, DNA topoisomerase (ATP-hydrolysing)
#2: DNA chain DNA (5'-D(*CP*AP*AP*AP*GP*GP*CP*GP*GP*TP*AP*AP*TP*AP*CP*GP*GP*TP*TP*AP*TP*CP*CP*AP*CP*AP*GP*AP*AP*TP*CP*AP*GP*G)-3')


Mass: 10525.804 Da / Num. of mol.: 2 / Source method: obtained synthetically / Details: V-site DNA / Source: (synth.) synthetic construct (others)
#3: DNA chain DNA (5'-D(*CP*CP*TP*GP*AP*TP*TP*CP*TP*GP*TP*GP*GP*AP*TP*AP*AP*CP*CP*GP*TP*AP*TP*TP*AP*CP*CP*GP*CP*CP*TP*TP*TP*G)-3')


Mass: 10391.674 Da / Num. of mol.: 2 / Source method: obtained synthetically / Details: V-site DNA / Source: (synth.) synthetic construct (others) / References: UniProt: Q59961*PLUS
#4: Chemical ChemComp-MG / MAGNESIUM ION


Mass: 24.305 Da / Num. of mol.: 3 / Source method: obtained synthetically / Formula: Mg
#5: Water ChemComp-HOH / water


Mass: 18.015 Da / Num. of mol.: 2 / Source method: isolated from a natural source / Formula: H2O
Has ligand of interestN
Has protein modificationN

-
Experimental details

-
Experiment

ExperimentMethod: X-RAY DIFFRACTION / Number of used crystals: 1

-
Sample preparation

CrystalDensity Matthews: 3.02 Å3/Da / Density % sol: 59.34 %
Crystal growTemperature: 296 K / Method: vapor diffusion, sitting drop / pH: 5
Details: 4% isopropanol, 50 mM sodium citrate, 600 nL + 400 nL (protein + precipitant) drop ratio, VAPOR DIFFUSION, SITTING DROP, temperature 296K

-
Data collection

DiffractionMean temperature: 100 K / Serial crystal experiment: N
Diffraction sourceSource: SYNCHROTRON / Site: Diamond / Beamline: I04-1 / Wavelength: 0.92 Å
DetectorType: DECTRIS PILATUS 2M / Detector: PIXEL / Date: Jul 23, 2012
RadiationMonochromator: Single bounce monochromator / Protocol: SINGLE WAVELENGTH / Monochromatic (M) / Laue (L): M / Scattering type: x-ray
Radiation wavelengthWavelength: 0.92 Å / Relative weight: 1
ReflectionResolution: 3.7→72.19 Å / Num. all: 39884 / Num. obs: 39884 / % possible obs: 99.75 % / Observed criterion σ(F): -1 / Observed criterion σ(I): -1 / Redundancy: 12.11 % / Rmerge(I) obs: 0.124 / Rsym value: 0.124 / Net I/σ(I): 3.1256
Reflection shellResolution: 3.7→3.9 Å / Redundancy: 11.46 % / Rmerge(I) obs: 0.45 / Mean I/σ(I) obs: 1.71 / Num. unique obs: 5609 / % possible all: 100

-
Processing

Software
NameVersionClassification
PHENIX1.21rc1_5058refinement
ISOLDE1.7.1model building
PHASERphasing
MOSFLMdata reduction
SCALAdata scaling
RefinementMethod to determine structure: MOLECULAR REPLACEMENT
Starting model: 3RAE,1EI1
Resolution: 3.7→71.82 Å / SU ML: 0.3403 / Cross valid method: FREE R-VALUE / σ(F): 1.4 / Phase error: 24.8969
Stereochemistry target values: GeoStd + Monomer Library + CDL v1.2
RfactorNum. reflection% reflection
Rfree0.277 2006 5.04 %
Rwork0.2297 37820 -
obs0.232 39826 99.69 %
Solvent computationShrinkage radii: 0.9 Å / VDW probe radii: 1.1 Å / Solvent model: FLAT BULK SOLVENT MODEL
Displacement parametersBiso mean: 95.05 Å2
Refinement stepCycle: LAST / Resolution: 3.7→71.82 Å
ProteinNucleic acidLigandSolventTotal
Num. atoms14889 1385 3 2 16279
Refine LS restraints
Refine-IDTypeDev idealNumber
X-RAY DIFFRACTIONf_bond_d0.002916682
X-RAY DIFFRACTIONf_angle_d0.618822990
X-RAY DIFFRACTIONf_chiral_restr0.04212692
X-RAY DIFFRACTIONf_plane_restr0.00462788
X-RAY DIFFRACTIONf_dihedral_angle_d14.87965809
LS refinement shell
Resolution (Å)Rfactor RfreeNum. reflection RfreeRfactor RworkNum. reflection RworkRefine-ID% reflection obs (%)
3.7-3.790.26091490.25422649X-RAY DIFFRACTION100
3.79-3.90.2591390.24282671X-RAY DIFFRACTION99.96
3.9-4.010.29531540.24342636X-RAY DIFFRACTION99.96
4.01-4.140.30131290.25362684X-RAY DIFFRACTION100
4.14-4.290.28211350.22232654X-RAY DIFFRACTION100
4.29-4.460.26081530.20432682X-RAY DIFFRACTION100
4.46-4.660.21561270.1922671X-RAY DIFFRACTION100
4.66-4.910.25151490.20932689X-RAY DIFFRACTION99.82
4.91-5.210.28391380.23812642X-RAY DIFFRACTION97.68
5.21-5.620.27621540.23412688X-RAY DIFFRACTION100
5.62-6.180.28821340.24422730X-RAY DIFFRACTION100
6.18-7.080.32181470.2412738X-RAY DIFFRACTION100
7.08-8.910.27611470.22042792X-RAY DIFFRACTION100
8.91-71.820.28611510.23522894X-RAY DIFFRACTION98.32

+
About Yorodumi

-
News

-
Feb 9, 2022. New format data for meta-information of EMDB entries

New format data for meta-information of EMDB entries

  • Version 3 of the EMDB header file is now the official format.
  • The previous official version 1.9 will be removed from the archive.

Related info.:EMDB header

External links:wwPDB to switch to version 3 of the EMDB data model

-
Aug 12, 2020. Covid-19 info

Covid-19 info

URL: https://pdbj.org/emnavi/covid19.php

New page: Covid-19 featured information page in EM Navigator.

Related info.:Covid-19 info / Mar 5, 2020. Novel coronavirus structure data

+
Mar 5, 2020. Novel coronavirus structure data

Novel coronavirus structure data

Related info.:Yorodumi Speices / Aug 12, 2020. Covid-19 info

External links:COVID-19 featured content - PDBj / Molecule of the Month (242):Coronavirus Proteases

+
Jan 31, 2019. EMDB accession codes are about to change! (news from PDBe EMDB page)

EMDB accession codes are about to change! (news from PDBe EMDB page)

  • The allocation of 4 digits for EMDB accession codes will soon come to an end. Whilst these codes will remain in use, new EMDB accession codes will include an additional digit and will expand incrementally as the available range of codes is exhausted. The current 4-digit format prefixed with “EMD-” (i.e. EMD-XXXX) will advance to a 5-digit format (i.e. EMD-XXXXX), and so on. It is currently estimated that the 4-digit codes will be depleted around Spring 2019, at which point the 5-digit format will come into force.
  • The EM Navigator/Yorodumi systems omit the EMD- prefix.

Related info.:Q: What is EMD? / ID/Accession-code notation in Yorodumi/EM Navigator

External links:EMDB Accession Codes are Changing Soon! / Contact to PDBj

+
Jul 12, 2017. Major update of PDB

Major update of PDB

  • wwPDB released updated PDB data conforming to the new PDBx/mmCIF dictionary.
  • This is a major update changing the version number from 4 to 5, and with Remediation, in which all the entries are updated.
  • In this update, many items about electron microscopy experimental information are reorganized (e.g. em_software).
  • Now, EM Navigator and Yorodumi are based on the updated data.

External links:wwPDB Remediation / Enriched Model Files Conforming to OneDep Data Standards Now Available in the PDB FTP Archive

-
Yorodumi

Thousand views of thousand structures

  • Yorodumi is a browser for structure data from EMDB, PDB, SASBDB, etc.
  • This page is also the successor to EM Navigator detail page, and also detail information page/front-end page for Omokage search.
  • The word "yorodu" (or yorozu) is an old Japanese word meaning "ten thousand". "mi" (miru) is to see.

Related info.:EMDB / PDB / SASBDB / Comparison of 3 databanks / Yorodumi Search / Aug 31, 2016. New EM Navigator & Yorodumi / Yorodumi Papers / Jmol/JSmol / Function and homology information / Changes in new EM Navigator and Yorodumi

Read more