Movie
 Controller


[English] 日本語
 Yorodumi
- PDB-1opk: Structural basis for the auto-inhibition of c-Abl tyrosine kinase -
+ Open data

- Basic information
| Entry | Database: PDB / ID: 1opk | ||||||
|---|---|---|---|---|---|---|---|
| Title | Structural basis for the auto-inhibition of c-Abl tyrosine kinase | ||||||
 Components | Proto-oncogene tyrosine-protein kinase ABL1 | ||||||
 Keywords | TRANSFERASE | ||||||
| Function / homology |  Function and homology information transitional one stage B cell differentiation / Role of ABL in ROBO-SLIT signaling / cerebellum morphogenesis / regulation of extracellular matrix organization / HDR through Single Strand Annealing (SSA) / B cell proliferation involved in immune response / B-1 B cell homeostasis / RHO GTPases Activate WASPs and WAVEs / neuroepithelial cell differentiation / positive regulation of Wnt signaling pathway, planar cell polarity pathway ...transitional one stage B cell differentiation / Role of ABL in ROBO-SLIT signaling / cerebellum morphogenesis / regulation of extracellular matrix organization / HDR through Single Strand Annealing (SSA) / B cell proliferation involved in immune response / B-1 B cell homeostasis / RHO GTPases Activate WASPs and WAVEs / neuroepithelial cell differentiation / positive regulation of Wnt signaling pathway, planar cell polarity pathway / alpha-beta T cell differentiation / microspike assembly / MLL4 and MLL3 complexes regulate expression of PPARG target genes in adipogenesis and hepatic steatosis / Turbulent (oscillatory, disturbed) flow shear stress activates signaling by PIEZO1 and integrins in endothelial cells / activated T cell proliferation / protein localization to cytoplasmic microtubule plus-end / DNA conformation change / regulation of cellular senescence / Cyclin D associated events in G1 / response to epinephrine / phospholipase C-inhibiting G protein-coupled receptor signaling pathway / negative regulation of ubiquitin-protein transferase activity / podocyte apoptotic process / Recruitment and ATM-mediated phosphorylation of repair and signaling proteins at DNA double strand breaks / Bergmann glial cell differentiation / regulation of postsynaptic specialization assembly / positive regulation of phospholipase C/protein kinase C signal transduction / regulation of modification of synaptic structure / delta-catenin binding / RUNX1 regulates transcription of genes involved in differentiation of HSCs / circulatory system development / positive regulation of extracellular matrix organization / Regulation of actin dynamics for phagocytic cup formation / neuropilin signaling pathway / neuropilin binding / Myogenesis / negative regulation of mitotic cell cycle / bubble DNA binding / spleen development / positive regulation of establishment of T cell polarity / neuromuscular process controlling balance / regulation of T cell differentiation / positive regulation of blood vessel branching / proline-rich region binding / post-embryonic development / platelet-derived growth factor receptor signaling pathway / positive regulation of dendrite development / B cell proliferation / regulation of Cdc42 protein signal transduction / negative regulation of cell-cell adhesion / syntaxin binding / negative regulation of cellular senescence / regulation of axon extension / mitogen-activated protein kinase binding / positive regulation of cell migration involved in sprouting angiogenesis / positive regulation of osteoblast proliferation / neural tube closure / thymus development / platelet-derived growth factor receptor-beta signaling pathway / cell leading edge / negative regulation of endothelial cell apoptotic process / myoblast proliferation / negative regulation of BMP signaling pathway / negative regulation of long-term synaptic potentiation / cardiac muscle cell proliferation / regulation of microtubule polymerization / associative learning / phagocytosis / ephrin receptor signaling pathway / cellular response to transforming growth factor beta stimulus / positive regulation of focal adhesion assembly / positive regulation of vasoconstriction / positive regulation of substrate adhesion-dependent cell spreading / endothelial cell migration / positive regulation of stress fiber assembly / substrate adhesion-dependent cell spreading / negative regulation of double-strand break repair via homologous recombination / positive regulation of T cell migration / ephrin receptor binding / positive regulation of mitotic cell cycle / four-way junction DNA binding / positive regulation of interleukin-2 production / ruffle / signal transduction in response to DNA damage / phosphotyrosine residue binding / actin filament polymerization / peptidyl-tyrosine phosphorylation / SH2 domain binding / positive regulation of endothelial cell migration / B cell receptor signaling pathway / integrin-mediated signaling pathway / response to endoplasmic reticulum stress / positive regulation of fibroblast proliferation / protein serine/threonine kinase activator activity / positive regulation of release of sequestered calcium ion into cytosol / regulation of actin cytoskeleton organization / protein kinase C binding / non-specific protein-tyrosine kinase / non-membrane spanning protein tyrosine kinase activity / positive regulation of type II interferon production Similarity search - Function | ||||||
| Biological species |   Mus musculus (house mouse) | ||||||
| Method |  X-RAY DIFFRACTION / SYNCHROTRON / MOLECULAR REPLACEMENT / Resolution: 1.8 Å | ||||||
 Authors | Nagar, B. / Hantschel, O. / Young, M.A. / Scheffzek, K. / Veach, D. / Bornmann, W. / Clarkson, B. / Superti-Furga, G. / Kuriyan, J. | ||||||
 Citation | Journal: Cell(Cambridge,Mass.) / Year: 2003 Title: Structural basis for the autoinhibition of c-Abl tyrosine kinase Authors: Nagar, B. / Hantschel, O. / Young, M.A. / Scheffzek, K. / Veach, D. / Bornmann, W. / Clarkson, B. / Superti-Furga, G. / Kuriyan, J. #1: Journal: Cell(Cambridge,Mass.) / Year: 2003Title: A myristoyl/phosphotyrosine switch regulates c-Abl Authors: Hantschel, O. / Nagar, B. / Guettler, S. / Kretzschmar, J. / Dorey, K. / Kuriyan, J. / Superti-Furga, G. | ||||||
| History |
| ||||||
| Remark 999 | SEQUENCE Numbering of the residues corresponds to the sequence database numbering of the isoform IV ...SEQUENCE Numbering of the residues corresponds to the sequence database numbering of the isoform IV of the protein. |
- Structure visualization
| Structure viewer | Molecule: MolmilJmol/JSmol |
|---|
- Downloads & links
-Download
| PDBx/mmCIF format | 1opk.cif.gz | 113.4 KB | Display | PDBx/mmCIF format |
|---|---|---|---|---|
| PDB format | pdb1opk.ent.gz | 84.3 KB | Display | PDB format |
| PDBx/mmJSON format | 1opk.json.gz | Tree view | PDBx/mmJSON format | |
| Others |  Other downloads |
-Validation report
| Arichive directory | https://data.pdbj.org/pub/pdb/validation_reports/op/1opkftp://data.pdbj.org/pub/pdb/validation_reports/op/1opk | HTTPS FTP |
|---|
-Related structure data
| Related structure data |  1opjC  1oplC  1m52S  2ablS C: citing same article ( S: Starting model for refinement |
|---|---|
| Similar structure data |

-Links
 PDBj
- Assembly
| Deposited unit | 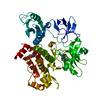
| ||||||||
|---|---|---|---|---|---|---|---|---|---|
| 1 |
| ||||||||
| Unit cell |
|
-Components
| #1: Protein | Mass: 56189.164 Da / Num. of mol.: 1 / Fragment: SH3-SH2-kinase domain / Mutation: D382N Source method: isolated from a genetically manipulated source Source: (gene. exp.) Mus musculus (house mouse) / Gene: ABL1 / Plasmid: PFASTBAC / Production host:   Spodoptera frugiperda (fall armyworm) / References: UniProt: P00520, EC: 2.7.1.112 |
|---|---|
| #2: Chemical | ChemComp-MYR /   Mass: 228.371 Da / Num. of mol.: 1 / Source method: obtained synthetically / Formula: C14H28O2 |
| #3: Chemical | ChemComp-P16 /   Mass: 427.283 Da / Num. of mol.: 1 / Source method: obtained synthetically / Formula: C21H16Cl2N4O2 |
| #4: Chemical | ChemComp-GOL /   Mass: 92.094 Da / Num. of mol.: 1 / Source method: obtained synthetically / Formula: C3H8O3 |
| #5: Water | ChemComp-HOH /  Mass: 18.015 Da / Num. of mol.: 270 / Source method: isolated from a natural source / Formula: H2O |
-Experimental details
-Experiment
| Experiment | Method: X-RAY DIFFRACTION / Number of used crystals: 1 |
|---|
- Sample preparation
| Crystal | Density Matthews: 2.49 Å3/Da / Density % sol: 50.22 % | |||||||||||||||||||||||||||||||||||||||||||||||||
|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|
| Crystal grow | Temperature: 293 K / Method: vapor diffusion, hanging drop / pH: 7 Details: 20% PEG 3350, 200 mM potassium nitrate, pH 7.0, VAPOR DIFFUSION, HANGING DROP, temperature 293K | |||||||||||||||||||||||||||||||||||||||||||||||||
| Crystal grow | *PLUS Temperature: 20 ℃ / pH: 8 | |||||||||||||||||||||||||||||||||||||||||||||||||
| Components of the solutions | *PLUS
|
-Data collection
| Diffraction | Mean temperature: 100 K |
|---|---|
| Diffraction source | Source: SYNCHROTRON / Site: ALS  / Beamline: 8.2.2 / Wavelength: 1.0781 Å |
| Detector | Type: ADSC QUANTUM 315 / Detector: CCD / Date: Dec 20, 2002 / Details: mirrors |
| Radiation | Monochromator: Double crystal Si(111) / Protocol: SINGLE WAVELENGTH / Monochromatic (M) / Laue (L): M / Scattering type: x-ray |
| Radiation wavelength | Wavelength: 1.0781 Å / Relative weight: 1 |
| Reflection | Resolution: 1.8→38 Å / Num. all: 50963 / Num. obs: 50963 / % possible obs: 99.9 % / Observed criterion σ(F): 0 / Observed criterion σ(I): -3 / Redundancy: 7.1 % / Biso Wilson estimate: 22 Å2 / Rsym value: 0.057 / Net I/σ(I): 29.6 |
| Reflection shell | Resolution: 1.8→1.86 Å / Redundancy: 6.7 % / Mean I/σ(I) obs: 3.5 / Num. unique all: 5078 / Rsym value: 0.584 / % possible all: 100 |
| Reflection | *PLUS Lowest resolution: 38 Å / Num. measured all: 560273 / Rmerge(I) obs: 0.057 |
| Reflection shell | *PLUS Highest resolution: 1.8 Å / % possible obs: 100 % / Rmerge(I) obs: 0.584 |
- Processing
| Software |
| ||||||||||||||||||||||||||||||||||||
|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|
| Refinement | Method to determine structure: MOLECULAR REPLACEMENT Starting model: PDB ENTRIES 1M52, 2ABL Resolution: 1.8→37.59 Å / Rfactor Rfree error: 0.004 / Isotropic thermal model: RESTRAINED / Cross valid method: THROUGHOUT / σ(F): 0 / Stereochemistry target values: Engh & Huber
| ||||||||||||||||||||||||||||||||||||
| Solvent computation | Solvent model: FLAT MODEL / Bsol: 35.0282 Å2 / ksol: 0.366818 e/Å3 | ||||||||||||||||||||||||||||||||||||
| Displacement parameters | Biso mean: 30.9 Å2
| ||||||||||||||||||||||||||||||||||||
| Refine analyze |
| ||||||||||||||||||||||||||||||||||||
| Refinement step | Cycle: LAST / Resolution: 1.8→37.59 Å
| ||||||||||||||||||||||||||||||||||||
| Refine LS restraints |
| ||||||||||||||||||||||||||||||||||||
| LS refinement shell | Resolution: 1.8→1.91 Å / Rfactor Rfree error: 0.013 / Total num. of bins used: 6
| ||||||||||||||||||||||||||||||||||||
| Xplor file |
| ||||||||||||||||||||||||||||||||||||
| Refinement | *PLUS Highest resolution: 1.8 Å / Lowest resolution: 38 Å | ||||||||||||||||||||||||||||||||||||
| Solvent computation | *PLUS | ||||||||||||||||||||||||||||||||||||
| Displacement parameters | *PLUS | ||||||||||||||||||||||||||||||||||||
| Refine LS restraints | *PLUS
|