Movie
 Controller


[English] 日本語
 Yorodumi
- PDB-45pc: 6-conformation model of Escherichia coli dihydrofolate reductase ... -
+ Open data

- Basic information
| Entry | Database: PDB / ID: 45pc | ||||||
|---|---|---|---|---|---|---|---|
| Title | 6-conformation model of Escherichia coli dihydrofolate reductase at 100K | ||||||
 Components | Dihydrofolate reductase | ||||||
 Keywords | OXIDOREDUCTASE / DHFR / folate reductase / tetrahydrofolate regulation | ||||||
| Function / homology |  Function and homology information methotrexate binding / dihydrofolic acid binding / response to methotrexate / 10-formyltetrahydrofolate biosynthetic process / folic acid biosynthetic process / folic acid binding / NADP+ binding / dihydrofolate metabolic process / dihydrofolate reductase / one-carbon metabolic process ...methotrexate binding / dihydrofolic acid binding / response to methotrexate / 10-formyltetrahydrofolate biosynthetic process / folic acid biosynthetic process / folic acid binding / NADP+ binding / dihydrofolate metabolic process / dihydrofolate reductase / one-carbon metabolic process / dihydrofolate reductase activity / folic acid metabolic process / NADPH binding / tetrahydrofolate biosynthetic process / NADP binding / response to antibiotic / response to xenobiotic stimulus / cytosol Similarity search - Function | ||||||
| Biological species |   Escherichia coli str. K-12 substr. MC4100 (bacteria) | ||||||
| Method |  X-RAY DIFFRACTION / SYNCHROTRON / MOLECULAR REPLACEMENT / Resolution: 0.85 Å | ||||||
 Authors | Passmore, S.K. | ||||||
| Funding support |  Australia, 1items
| ||||||
 Citation | Journal: Biorxiv / Year: 2026 Title: Optimizing the connectivity of protein conformations to untangle ensemble refinement Authors: Passmore, S.K. / Holton, J.M. / Zatsepin, N.A. / Martin, A.V. #1: Journal: Structure / Year: 2014Title: Crystal cryocooling distorts conformational heterogeneity in a model Michaelis complex of DHFR. Authors: Keedy, D.A. / van den Bedem, H. / Sivak, D.A. / Petsko, G.A. / Ringe, D. / Wilson, M.A. / Fraser, J.S. | ||||||
| History |
| ||||||
| Remark 0 | THIS ENTRY 45PC REFLECTS AN ALTERNATIVE MODELING OF THE ORIGINAL DATA IN 4PSS, DETERMINED BY D.A. ...THIS ENTRY 45PC REFLECTS AN ALTERNATIVE MODELING OF THE ORIGINAL DATA IN 4PSS, DETERMINED BY D.A.Keedy, H.van den Bedem, D.A.Sivak, G.A.Petsko, D.Ringe, M.A.Wilson, J.S.Fraser. |
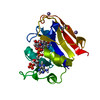
- Structure visualization
| Structure viewer | Molecule: MolmilJmol/JSmol |
|---|
- Downloads & links
-Download
| PDBx/mmCIF format | 45pc.cif.gz | 485.4 KB | Display | PDBx/mmCIF format |
|---|---|---|---|---|
| PDB format | pdb45pc.ent.gz | 343.1 KB | Display | PDB format |
| PDBx/mmJSON format | 45pc.json.gz | Tree view | PDBx/mmJSON format | |
| Others |  Other downloads |
-Validation report
| Arichive directory | https://data.pdbj.org/pub/pdb/validation_reports/5p/45pcftp://data.pdbj.org/pub/pdb/validation_reports/5p/45pc | HTTPS FTP |
|---|
-Related structure data


-Links
 PDBj
- Assembly
| Deposited unit | 
| ||||||||||||
|---|---|---|---|---|---|---|---|---|---|---|---|---|---|
| 1 |
| ||||||||||||
| Unit cell |
|
-Components
-Protein , 1 types, 1 molecules A
| #1: Protein | Mass: 18051.338 Da / Num. of mol.: 1 Source method: isolated from a genetically manipulated source Source: (gene. exp.) Escherichia coli str. K-12 substr. MC4100 (bacteria)Gene: folA, tmrA, b0048, JW0047 / Production host: Escherichia coli (E. coli) / References: UniProt: P0ABQ4, dihydrofolate reductase |
|---|
-Non-polymers , 5 types, 2666 molecules 
| #2: Chemical | ChemComp-FOL /  Mass: 441.397 Da / Num. of mol.: 1 / Source method: obtained synthetically / Formula: C19H19N7O6 | ||||
|---|---|---|---|---|---|
| #3: Chemical | ChemComp-NAP /  Mass: 743.405 Da / Num. of mol.: 1 / Source method: obtained synthetically / Formula: C21H28N7O17P3 | ||||
| #4: Chemical |  Mass: 54.938 Da / Num. of mol.: 2 / Source method: obtained synthetically / Formula: Mn #5: Chemical | ChemComp-CL / |  Mass: 35.453 Da / Num. of mol.: 1 / Source method: obtained synthetically / Formula: Cl #6: Water | ChemComp-HOH / | Mass: 18.015 Da / Num. of mol.: 2661 / Source method: isolated from a natural source / Formula: H2O |
-Details
| Has ligand of interest | N |
|---|---|
| Has protein modification | Y |
-Experimental details
-Experiment
| Experiment | Method: X-RAY DIFFRACTION / Number of used crystals: 1 |
|---|
- Sample preparation
| Crystal | Density Matthews: 2.07 Å3/Da / Density % sol: 40.62 % / Description: author used sf file from pdb entry 4PSS |
|---|---|
| Crystal grow | Temperature: 277 K / Method: vapor diffusion, hanging drop / pH: 7 Details: 17% PEG 400, 20 mM imidazole pH 7.0, 125 mM MnCl2, VAPOR DIFFUSION, HANGING DROP, temperature 277K |
-Data collection
| Diffraction | Mean temperature: 100 K / Serial crystal experiment: N |
|---|---|
| Diffraction source | Source: SYNCHROTRON / Site: SSRL  / Beamline: BL11-1 / Wavelength: 0.9 Å |
| Detector | Type: ADSC QUANTUM 315 / Detector: CCD / Date: Apr 25, 2005 |
| Radiation | Protocol: SINGLE WAVELENGTH / Monochromatic (M) / Laue (L): M / Scattering type: x-ray |
| Radiation wavelength | Wavelength: 0.9 Å / Relative weight: 1 |
| Reflection | Resolution: 0.849→40.78 Å / Num. obs: 130200 / % possible obs: 98.2 % / Redundancy: 5.5 % / Biso Wilson estimate: 7.68 Å2 / Rsym value: 0.049 / Net I/σ(I): 36.3 |
| Reflection shell | Resolution: 0.85→0.88 Å / Redundancy: 4 % / Rmerge(I) obs: 0.55 / Mean I/σ(I) obs: 2.3 / Num. unique obs: 12585 / % possible all: 97.2 |
- Processing
| Software |
| |||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||
|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|
| Refinement | Method to determine structure: MOLECULAR REPLACEMENT / Resolution: 0.85→40.78 Å / SU ML: 0.0695 / Cross valid method: FREE R-VALUE / σ(F): 1.34 / Phase error: 7.8622 Stereochemistry target values: GeoStd + Monomer Library + CDL v1.2
| |||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||
| Solvent computation | Shrinkage radii: 0.9 Å / VDW probe radii: 1.1 Å / Solvent model: FLAT BULK SOLVENT MODEL | |||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||
| Displacement parameters | Biso mean: 7.47 Å2 | |||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||
| Refinement step | Cycle: LAST / Resolution: 0.85→40.78 Å
| |||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||
| Refine LS restraints |
| |||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||
| LS refinement shell |
|