Movie
 Controller


+ Open data

- Basic information
| Entry | Database: PDB / ID: 45kk | ||||||
|---|---|---|---|---|---|---|---|
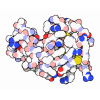
| Title | 6-conformation model of Hen Egg White Lysozyme at 100K | ||||||
 Components | Lysozyme C | ||||||
 Keywords | ISOMERASE / lysozyme | ||||||
| Function / homology |  Function and homology information Lactose synthesis / Antimicrobial peptides / Neutrophil degranulation / beta-N-acetylglucosaminidase activity / cell wall macromolecule catabolic process / lysozyme / lysozyme activity / killing of cells of another organism / defense response to Gram-negative bacterium / defense response to bacterium ...Lactose synthesis / Antimicrobial peptides / Neutrophil degranulation / beta-N-acetylglucosaminidase activity / cell wall macromolecule catabolic process / lysozyme / lysozyme activity / killing of cells of another organism / defense response to Gram-negative bacterium / defense response to bacterium / defense response to Gram-positive bacterium / Golgi apparatus / endoplasmic reticulum / extracellular region / identical protein binding / cytoplasm Similarity search - Function | ||||||
| Biological species |   Gallus gallus (chicken) | ||||||
| Method |  X-RAY DIFFRACTION / SYNCHROTRON / MOLECULAR REPLACEMENT / Resolution: 1.2 Å | ||||||
 Authors | Passmore, S.K. | ||||||
| Funding support |  Australia, 1items
| ||||||
 Citation | Journal: Biorxiv / Year: 2026 Title: Optimizing the connectivity of protein conformations to untangle ensemble refinement Authors: Passmore, S.K. / Holton, J.M. / Zatsepin, N.A. / Martin, A.V. #1: Journal: J.Synchrotron Radiat. / Year: 2017Title: Conformational variation of proteins at room temperature is not dominated by radiation damage. Authors: Russi, S. / Gonzalez, A. / Kenner, L.R. / Keedy, D.A. / Fraser, J.S. / van den Bedem, H. | ||||||
| History |
| ||||||
| Remark 0 | THIS ENTRY 45KK REFLECTS AN ALTERNATIVE MODELING OF THE ORIGINAL DATA IN 5KXK, DETERMINED BY S. ...THIS ENTRY 45KK REFLECTS AN ALTERNATIVE MODELING OF THE ORIGINAL DATA IN 5KXK, DETERMINED BY S.Russi, A.Gonzalez, L.R.Kenner, D.A.Keedy, J.S.Fraser, H.van den Bedem. |
- Structure visualization
| Structure viewer | Molecule: MolmilJmol/JSmol |
|---|
- Downloads & links
-Download
| PDBx/mmCIF format | 45kk.cif.gz | 308.4 KB | Display | PDBx/mmCIF format |
|---|---|---|---|---|
| PDB format | pdb45kk.ent.gz | 243.5 KB | Display | PDB format |
| PDBx/mmJSON format | 45kk.json.gz | Tree view | PDBx/mmJSON format | |
| Others |  Other downloads |
-Validation report
| Arichive directory | https://data.pdbj.org/pub/pdb/validation_reports/5k/45kkftp://data.pdbj.org/pub/pdb/validation_reports/5k/45kk | HTTPS FTP |
|---|
-Related structure data


-Links
 PDBj
- Assembly
| Deposited unit | 
| ||||||||||||||||||||||||||||||||||||||||||
|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|
| 1 |
| ||||||||||||||||||||||||||||||||||||||||||
| Unit cell |
| ||||||||||||||||||||||||||||||||||||||||||
| Components on special symmetry positions |
|

-Components
| #1: Protein | Mass: 14331.160 Da / Num. of mol.: 1 Source method: isolated from a genetically manipulated source Source: (gene. exp.) Gallus gallus (chicken) / Gene: LYZ / Production host: Gallus gallus (chicken) / References: UniProt: P00698, lysozyme | ||||||||
|---|---|---|---|---|---|---|---|---|---|
| #2: Chemical | ChemComp-ACT /   Mass: 59.044 Da / Num. of mol.: 1 / Source method: obtained synthetically / Formula: C2H3O2 | ||||||||
| #3: Chemical |   Mass: 22.990 Da / Num. of mol.: 2 / Source method: obtained synthetically / Formula: Na #4: Chemical |   Mass: 35.453 Da / Num. of mol.: 3 / Source method: obtained synthetically / Formula: Cl #5: Water | ChemComp-HOH / |  Mass: 18.015 Da / Num. of mol.: 1468 / Source method: isolated from a natural source / Formula: H2O Has ligand of interest | N | Has protein modification | Y | |
-Experimental details
-Experiment
| Experiment | Method: X-RAY DIFFRACTION / Number of used crystals: 1 |
|---|
- Sample preparation
| Crystal | Density Matthews: 1.94 Å3/Da / Density % sol: 36.55 % / Description: AUTHOR USED THE SF DATA FROM ENTRY 5KXK |
|---|---|
| Crystal grow | Temperature: 291 K / Method: vapor diffusion, sitting drop / pH: 4.5 Details: CRYSTALS WERE GROWN BY MIXING EQUAL VOLUMES OF WELL SOLUTION (1.0 M SODIUM CHLORIDE, 50 MM SODIUM ACETATE PH 4.5) AND PROTEIN (40-60 MG/ML) |
-Data collection
| Diffraction | Mean temperature: 100 K / Serial crystal experiment: N |
|---|---|
| Diffraction source | Source: SYNCHROTRON / Site: SSRL  / Beamline: BL14-1 / Wavelength: 1.21549 Å |
| Detector | Type: MARMOSAIC 325 mm CCD / Detector: CCD / Date: Jul 19, 2013 |
| Radiation | Monochromator: M / Protocol: SINGLE WAVELENGTH / Monochromatic (M) / Laue (L): M / Scattering type: x-ray |
| Radiation wavelength | Wavelength: 1.21549 Å / Relative weight: 1 |
| Reflection | Resolution: 1.198→38.65 Å / Num. obs: 35966 / % possible obs: 99.6 % / Redundancy: 6.6 % / Biso Wilson estimate: 9.29 Å2 / Rsym value: 0.025 / Net I/σ(I): 45.8 |
| Reflection shell | Resolution: 1.2→1.26 Å / Redundancy: 5.8 % / Rmerge(I) obs: 0.098 / Mean I/σ(I) obs: 8 / Num. unique obs: 4943 / % possible all: 97.7 |
- Processing
| Software |
| |||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||
|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|
| Refinement | Method to determine structure: MOLECULAR REPLACEMENT / Resolution: 1.2→38.64 Å / SU ML: 0.0477 / Cross valid method: FREE R-VALUE / σ(F): 1.36 / Phase error: 5.7449 Stereochemistry target values: GeoStd + Monomer Library + CDL v1.2 Details: Model was refined over loops consisting of a single unrestrained refinement macro-cycle, connectivity optimization (reassigning altloc labels), then 10+ restrained refinement macro-cycles. ...Details: Model was refined over loops consisting of a single unrestrained refinement macro-cycle, connectivity optimization (reassigning altloc labels), then 10+ restrained refinement macro-cycles. Conformation-aware water picking and manual adjustments were also performed in later loops.
| |||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||
| Solvent computation | Shrinkage radii: 0.9 Å / VDW probe radii: 1.1 Å / Solvent model: FLAT BULK SOLVENT MODEL | |||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||
| Displacement parameters | Biso mean: 8.93 Å2 | |||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||
| Refinement step | Cycle: LAST / Resolution: 1.2→38.64 Å
| |||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||
| Refine LS restraints |
| |||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||
| LS refinement shell |
|