SEQUENCE THE CONSTRUCT WAS EXPRESSED WITH A PURIFICATION TAG MGSDKIHHHHHHENLYFQG. THE TAG WAS ... SEQUENCE THE CONSTRUCT WAS EXPRESSED WITH A PURIFICATION TAG MGSDKIHHHHHHENLYFQG. THE TAG WAS REMOVED WITH TEV PROTEASE LEAVING ONLY A GLYCINE (0) FOLLOWED BY THE TARGET SEQUENCE.
Type: MARMOSAIC 325 mm CCD / Detector: CCD / Date: Nov 20, 2007 / Details: Flat mirror (vertical focusing)
Radiation
Monochromator: Single crystal Si(111) bent (horizontal focusing) Protocol: SINGLE WAVELENGTH / Monochromatic (M) / Laue (L): M / Scattering type: x-ray
Radiation wavelength
Wavelength: 0.97916 Å / Relative weight: 1
Reflection
Resolution: 1.85→40.825 Å / Num. obs: 45283 / % possible obs: 99.5 % / Redundancy: 5.2 % / Biso Wilson estimate: 13.78 Å2 / Rmerge(I) obs: 0.146 / Rsym value: 0.146 / Net I/σ(I): 5.1
Reflection shell
Diffraction-ID: 1
Resolution (Å)
Redundancy (%)
Rmerge(I) obs
Mean I/σ(I) obs
Num. measured all
Num. unique all
Rsym value
% possible all
1.85-1.95
4.6
0.611
1.3
29250
6400
0.611
97.2
1.95-2.07
5.3
0.466
1.7
32802
6200
0.466
99.7
2.07-2.21
5.3
0.353
2.2
31028
5868
0.353
99.8
2.21-2.39
5.3
0.272
2.9
28978
5465
0.272
99.8
2.39-2.62
5.3
0.219
3.5
26963
5081
0.219
99.9
2.62-2.93
5.3
0.148
5.2
24120
4548
0.148
99.9
2.93-3.38
5.3
0.094
8.1
21467
4049
0.094
100
3.38-4.14
5.3
0.062
11.5
18146
3426
0.062
100
4.14-5.85
5.2
0.055
12.5
14103
2722
0.055
100
5.85-40.825
4.9
0.046
13.8
7489
1524
0.046
99.8
-
Phasing
Phasing
Method: SAD
-
Processing
Software
Name
Version
Classification
NB
REFMAC
5.4.0062
refinement
PHENIX
refinement
SHELX
phasing
MolProbity
3beta29
modelbuilding
SCALA
datascaling
PDB_EXTRACT
3
dataextraction
MAR345
CCD
datacollection
XDS
datareduction
SHARP
phasing
SHELXD
phasing
Refinement
Method to determine structure: SAD / Resolution: 1.85→40.825 Å / Cor.coef. Fo:Fc: 0.95 / Cor.coef. Fo:Fc free: 0.911 / SU B: 7.961 / SU ML: 0.123 / TLS residual ADP flag: LIKELY RESIDUAL / Cross valid method: THROUGHOUT / σ(F): 0 / ESU R: 0.154 / ESU R Free: 0.151 Stereochemistry target values: MAXIMUM LIKELIHOOD WITH PHASES Details: 1. HYDROGENS HAVE BEEN ADDED IN THE RIDING POSITIONS. 2. A MET-INHIBITION PROTOCOL WAS USED FOR SELENOMETHIONINE INCORPORATION DURING PROTEIN EXPRESSION. THE OCCUPANCY OF THE SE ATOMS IN THE ...Details: 1. HYDROGENS HAVE BEEN ADDED IN THE RIDING POSITIONS. 2. A MET-INHIBITION PROTOCOL WAS USED FOR SELENOMETHIONINE INCORPORATION DURING PROTEIN EXPRESSION. THE OCCUPANCY OF THE SE ATOMS IN THE MSE RESIDUES WAS REDUCED TO 0.75 FOR THE REDUCED SCATTERING POWER DUE TO PARTIAL S-MET INCORPORATION. 3. ATOM RECORD CONTAINS RESIDUAL B FACTORS ONLY. 4. RAMACHANDRAN OUTLIER A115G IS SUPPORTED BY CLEARLY DEFINED ELECTRON DENSITY.
Rfactor
Num. reflection
% reflection
Selection details
Rfree
0.242
2285
5 %
RANDOM
Rwork
0.185
-
-
-
obs
0.188
45282
99.41 %
-
Solvent computation
Ion probe radii: 0.8 Å / Shrinkage radii: 0.8 Å / VDW probe radii: 1.2 Å / Solvent model: MASK
Displacement parameters
Biso mean: 14.309 Å2
Baniso -1
Baniso -2
Baniso -3
1-
0.21 Å2
0 Å2
-1 Å2
2-
-
0.64 Å2
0 Å2
3-
-
-
-1.33 Å2
Refinement step
Cycle: LAST / Resolution: 1.85→40.825 Å
Protein
Nucleic acid
Ligand
Solvent
Total
Num. atoms
4152
0
1
492
4645
Refine LS restraints
Refine-ID
Type
Dev ideal
Dev ideal target
Number
X-RAY DIFFRACTION
r_bond_refined_d
0.016
0.021
4314
X-RAY DIFFRACTION
r_bond_other_d
0.001
0.02
2717
X-RAY DIFFRACTION
r_angle_refined_deg
1.488
1.943
5921
X-RAY DIFFRACTION
r_angle_other_deg
1.011
3
6679
X-RAY DIFFRACTION
r_dihedral_angle_1_deg
5.831
5
567
X-RAY DIFFRACTION
r_dihedral_angle_2_deg
43.209
24.497
189
X-RAY DIFFRACTION
r_dihedral_angle_3_deg
14.089
15
665
X-RAY DIFFRACTION
r_dihedral_angle_4_deg
16.748
15
23
X-RAY DIFFRACTION
r_chiral_restr
0.089
0.2
702
X-RAY DIFFRACTION
r_gen_planes_refined
0.007
0.021
4904
X-RAY DIFFRACTION
r_gen_planes_other
0.001
0.02
835
X-RAY DIFFRACTION
r_mcbond_it
0.797
1.5
2759
X-RAY DIFFRACTION
r_mcbond_other
0.213
1.5
1119
X-RAY DIFFRACTION
r_mcangle_it
1.369
2
4433
X-RAY DIFFRACTION
r_scbond_it
2.338
3
1555
X-RAY DIFFRACTION
r_scangle_it
3.457
4.5
1477
LS refinement shell
Resolution: 1.85→1.898 Å / Total num. of bins used: 20
Rfactor
Num. reflection
% reflection
Rfree
0.339
165
-
Rwork
0.294
2985
-
all
-
3150
-
obs
-
-
94.48 %
Refinement TLS params.
Method: refined / Refine-ID: X-RAY DIFFRACTION
ID
L11 (°2)
L12 (°2)
L13 (°2)
L22 (°2)
L23 (°2)
L33 (°2)
S11 (Å °)
S12 (Å °)
S13 (Å °)
S21 (Å °)
S22 (Å °)
S23 (Å °)
S31 (Å °)
S32 (Å °)
S33 (Å °)
T11 (Å2)
T12 (Å2)
T13 (Å2)
T22 (Å2)
T23 (Å2)
T33 (Å2)
Origin x (Å)
Origin y (Å)
Origin z (Å)
1
0.4885
0.0803
0.3685
0.3325
0.1066
1.463
-0.0212
0.0185
-0.002
-0.0023
0.0259
-0.0244
-0.0281
0.0078
-0.0047
-0.0489
-0.0061
0.0036
0.001
-0.004
-0.0302
27.0845
1.6857
7.9176
2
0.4153
0.1138
0.5663
0.0538
0.2503
1.1729
-0.0576
-0.0487
0.0481
0.0202
0.0028
-0.0128
-0.0275
-0.112
0.0547
-0.0342
-0.0049
-0.0029
0.0017
-0.0038
-0.0159
29.5191
5.1165
28.3432
3
0.368
0.0358
-0.0661
0.6955
-0.0043
0.984
0.0111
0.026
-0.0307
0.0493
-0.0209
-0.018
0.0477
0.0086
0.0098
-0.0094
-0.0128
-0.0045
-0.0477
0.0027
-0.0133
50.5933
2.4252
65.9308
4
0.638
0.2936
0.6545
0.6743
0.2025
0.9638
-0.0112
-0.0668
0.0285
-0.0676
-0.0186
0.064
-0.0484
-0.0794
0.0297
-0.0068
-0.0133
0.0071
-0.0315
-0.0046
-0.0213
42.4692
7.9142
47.5341
Refinement TLS group
ID
Refine-ID
Refine TLS-ID
Auth asym-ID
Label asym-ID
Auth seq-ID
Label seq-ID
1
X-RAY DIFFRACTION
1
A
A
1 - 130
2 - 131
2
X-RAY DIFFRACTION
2
A
A
131 - 274
132 - 275
3
X-RAY DIFFRACTION
3
B
B
1 - 130
2 - 131
4
X-RAY DIFFRACTION
4
B
B
131 - 274
132 - 275
+
About Yorodumi
-
News
-
Feb 9, 2022. New format data for meta-information of EMDB entries
New format data for meta-information of EMDB entries
Version 3 of the EMDB header file is now the official format.
The previous official version 1.9 will be removed from the archive.
In the structure databanks used in Yorodumi, some data are registered as the other names, "COVID-19 virus" and "2019-nCoV". Here are the details of the virus and the list of structure data.
Jan 31, 2019. EMDB accession codes are about to change! (news from PDBe EMDB page)
EMDB accession codes are about to change! (news from PDBe EMDB page)
The allocation of 4 digits for EMDB accession codes will soon come to an end. Whilst these codes will remain in use, new EMDB accession codes will include an additional digit and will expand incrementally as the available range of codes is exhausted. The current 4-digit format prefixed with “EMD-” (i.e. EMD-XXXX) will advance to a 5-digit format (i.e. EMD-XXXXX), and so on. It is currently estimated that the 4-digit codes will be depleted around Spring 2019, at which point the 5-digit format will come into force.
The EM Navigator/Yorodumi systems omit the EMD- prefix.
Related info.:Q: What is EMD? / ID/Accession-code notation in Yorodumi/EM Navigator
Yorodumi is a browser for structure data from EMDB, PDB, SASBDB, etc.
This page is also the successor to EM Navigator detail page, and also detail information page/front-end page for Omokage search.
The word "yorodu" (or yorozu) is an old Japanese word meaning "ten thousand". "mi" (miru) is to see.
Related info.:EMDB / PDB / SASBDB / Comparison of 3 databanks / Yorodumi Search / Aug 31, 2016. New EM Navigator & Yorodumi / Yorodumi Papers / Jmol/JSmol / Function and homology information / Changes in new EM Navigator and Yorodumi
 Movie
 Controller


 Yorodumi
 Open data

 Basic information
 Components
 Keywords
 Function and homology information
 Lactobacillus casei (bacteria)
 X-RAY DIFFRACTION /
 Authors
 Citation
 Structure visualization
 Downloads & links
 Other downloads

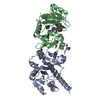
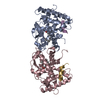
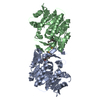
 PDBj
 Assembly
 Escherichia coli (E. coli) / Strain (production host): HK100 / References: UniProt: Q034N3
 Mass: 35.453 Da / Num. of mol.: 1 / Source method: obtained synthetically / Formula: Cl
 Mass: 18.015 Da / Num. of mol.: 492 / Source method: isolated from a natural source / Formula: H2O
 Sample preparation
 / Beamline: BL11-1 / Wavelength: 0.97916 Å
 Processing