Movie
 Controller


[English] 日本語
 Yorodumi
- PDB-29nx: X-ray structure of the adduct formed upon reaction of Lysozyme wi... -
+ Open data

- Basic information
| Entry | Database: PDB / ID: 29nx | ||||||
|---|---|---|---|---|---|---|---|
| Title | X-ray structure of the adduct formed upon reaction of Lysozyme with [Ru2Cl(D-p-TolF)(O2CCH3)3] | ||||||
 Components | Lysozyme C | ||||||
 Keywords | METAL BINDING PROTEIN / ruthenium / diruthenium / protein interaction / metallodrug | ||||||
| Function / homology |  Function and homology information Lactose synthesis / Antimicrobial peptides / Neutrophil degranulation / beta-N-acetylglucosaminidase activity / cell wall macromolecule catabolic process / lysozyme / lysozyme activity / killing of cells of another organism / defense response to Gram-negative bacterium / defense response to bacterium ...Lactose synthesis / Antimicrobial peptides / Neutrophil degranulation / beta-N-acetylglucosaminidase activity / cell wall macromolecule catabolic process / lysozyme / lysozyme activity / killing of cells of another organism / defense response to Gram-negative bacterium / defense response to bacterium / defense response to Gram-positive bacterium / Golgi apparatus / endoplasmic reticulum / : / identical protein binding / cytoplasm Similarity search - Function | ||||||
| Biological species |   Gallus gallus (chicken) | ||||||
| Method |  X-RAY DIFFRACTION / SYNCHROTRON / MOLECULAR REPLACEMENT / Resolution: 1.3 Å | ||||||
 Authors | Teran, A. / Ferraro, G. / Merlino, A. | ||||||
| Funding support | 1items
| ||||||
 Citation | Journal: Inorg.Chem. / Year: 2026 Title: Protein Recognition and Amyloid Remodeling Governed by Paddlewheel Diruthenium Coordination Chemistry. Authors: La Manna, S. / Florio, D. / Ferraro, G. / Policino, M.R. / Altieri, F. / Teran, A. / Herrero, S. / Merlino, A. / Marasco, D. | ||||||
| History |
|
- Structure visualization
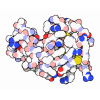
| Structure viewer | Molecule: MolmilJmol/JSmol |
|---|
- Downloads & links
-Download
| PDBx/mmCIF format | 29nx.cif.gz | 50.5 KB | Display | PDBx/mmCIF format |
|---|---|---|---|---|
| PDB format | pdb29nx.ent.gz | Display | PDB format | |
| PDBx/mmJSON format | 29nx.json.gz | Tree view | PDBx/mmJSON format | |
| Others |  Other downloads |
-Validation report
| Arichive directory | https://data.pdbj.org/pub/pdb/validation_reports/9n/29nxftp://data.pdbj.org/pub/pdb/validation_reports/9n/29nx | HTTPS FTP |
|---|
-Related structure data


-Links
 PDBj
- Assembly
| Deposited unit | 
| ||||||||||||||||||
|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|
| 1 |
| ||||||||||||||||||
| Unit cell |
| ||||||||||||||||||
| Components on special symmetry positions |
|
-Components
-Protein , 1 types, 1 molecules A
| #1: Protein | Mass: 14331.160 Da / Num. of mol.: 1 / Source method: isolated from a natural source / Source: (natural) Gallus gallus (chicken) / References: UniProt: P00698, lysozyme |
|---|
-Non-polymers , 5 types, 113 molecules 
| #2: Chemical | ChemComp-CL /  Mass: 35.453 Da / Num. of mol.: 1 / Source method: obtained synthetically / Formula: Cl | ||||
|---|---|---|---|---|---|
| #3: Chemical | ChemComp-NA /  Mass: 22.990 Da / Num. of mol.: 1 / Source method: obtained synthetically / Formula: Na | ||||
| #4: Chemical | ChemComp-NO3 /  Mass: 62.005 Da / Num. of mol.: 5 / Source method: obtained synthetically / Formula: NO3 #5: Chemical | Mass: 605.589 Da / Num. of mol.: 3 / Source method: obtained synthetically / Formula: C21H27N2O6Ru2 / Feature type: SUBJECT OF INVESTIGATION #6: Water | ChemComp-HOH / | Mass: 18.015 Da / Num. of mol.: 103 / Source method: isolated from a natural source / Formula: H2O |
-Details
| Has ligand of interest | Y |
|---|---|
| Has protein modification | Y |
-Experimental details
-Experiment
| Experiment | Method: X-RAY DIFFRACTION / Number of used crystals: 1 |
|---|
- Sample preparation
| Crystal | Density Matthews: 1.97 Å3/Da / Density % sol: 37.52 % |
|---|---|
| Crystal grow | Temperature: 293 K / Method: vapor diffusion, hanging drop / pH: 4 Details: 20% ethylene glycol, 0.1 M sodium acetate at pH 4.0, and 0.6 M sodium nitrate |
-Data collection
| Diffraction | Mean temperature: 100 K / Serial crystal experiment: N |
|---|---|
| Diffraction source | Source: SYNCHROTRON / Site: ELETTRA  / Beamline: 11.2C / Wavelength: 1 Å |
| Detector | Type: DECTRIS PILATUS 6M / Detector: PIXEL / Date: Jun 17, 2022 |
| Radiation | Protocol: SINGLE WAVELENGTH / Monochromatic (M) / Laue (L): M / Scattering type: x-ray |
| Radiation wavelength | Wavelength: 1 Å / Relative weight: 1 |
| Reflection | Resolution: 1.3→54.822 Å / Num. obs: 28706 / % possible obs: 99.9 % / Redundancy: 12.9 % / CC1/2: 1 / Net I/σ(I): 31.5 |
| Reflection shell | Resolution: 1.3→1.32 Å / Num. unique obs: 1400 / CC1/2: 0.775 |
- Processing
| Software |
| |||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||
|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|
| Refinement | Method to determine structure: MOLECULAR REPLACEMENT / Resolution: 1.3→54.822 Å / Cor.coef. Fo:Fc: 0.958 / Cor.coef. Fo:Fc free: 0.956 / SU B: 1.118 / SU ML: 0.047 / Cross valid method: THROUGHOUT / ESU R: 0.063 / ESU R Free: 0.065 Details: Hydrogens have been added in their riding positions
| |||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||
| Solvent computation | Ion probe radii: 0.8 Å / Shrinkage radii: 0.8 Å / VDW probe radii: 1.2 Å / Solvent model: MASK BULK SOLVENT | |||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||
| Displacement parameters | Biso mean: 22.865 Å2
| |||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||
| Refinement step | Cycle: LAST / Resolution: 1.3→54.822 Å
| |||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||
| Refine LS restraints |
| |||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||
| LS refinement shell | Refine-ID: X-RAY DIFFRACTION / Total num. of bins used: 20
|