Mass: 18.015 Da / Num. of mol.: 126 / Source method: isolated from a natural source / Formula: H2O
Has ligand of interest
Y
Has protein modification
Y
-
Experimental details
-
Experiment
Experiment
Method: X-RAY DIFFRACTION / Number of used crystals: 1
-
Sample preparation
Crystal
Density Matthews: 1.97 Å3/Da / Density % sol: 37.42 %
Crystal grow
Temperature: 293 K / Method: vapor diffusion, hanging drop / pH: 4 Details: 20% ethylene glycol, 0.1 M sodium acetate at pH 4.0, and 0.6 M sodium nitrate
-
Data collection
Diffraction
Mean temperature: 100 K / Serial crystal experiment: N
Method to determine structure: MOLECULAR REPLACEMENT / Resolution: 1.42→38.945 Å / Cor.coef. Fo:Fc: 0.963 / Cor.coef. Fo:Fc free: 0.952 / SU B: 1.455 / SU ML: 0.057 / Cross valid method: THROUGHOUT / ESU R: 0.079 / ESU R Free: 0.08 Details: Hydrogens have been added in their riding positions
Rfactor
Num. reflection
% reflection
Selection details
Rfree
0.2255
1167
5.273 %
RANDOM
Rwork
0.1927
20965
-
-
all
0.194
-
-
-
obs
-
22132
99.896 %
-
Solvent computation
Ion probe radii: 0.8 Å / Shrinkage radii: 0.8 Å / VDW probe radii: 1.2 Å / Solvent model: MASK BULK SOLVENT
Displacement parameters
Biso mean: 22.678 Å2
Baniso -1
Baniso -2
Baniso -3
1-
-0.057 Å2
-0 Å2
0 Å2
2-
-
-0.057 Å2
-0 Å2
3-
-
-
0.114 Å2
Refinement step
Cycle: LAST / Resolution: 1.42→38.945 Å
Protein
Nucleic acid
Ligand
Solvent
Total
Num. atoms
1001
0
50
126
1177
Refine LS restraints
Refine-ID
Type
Dev ideal
Dev ideal target
Number
X-RAY DIFFRACTION
r_bond_refined_d
0.01
0.012
1166
X-RAY DIFFRACTION
r_bond_other_d
0.001
0.016
1026
X-RAY DIFFRACTION
r_angle_refined_deg
2.024
1.833
1597
X-RAY DIFFRACTION
r_angle_other_deg
1.51
1.814
2347
X-RAY DIFFRACTION
r_dihedral_angle_1_deg
6.396
5
144
X-RAY DIFFRACTION
r_dihedral_angle_2_deg
10.555
5
14
X-RAY DIFFRACTION
r_dihedral_angle_3_deg
12.965
10
189
X-RAY DIFFRACTION
r_dihedral_angle_6_deg
14.264
10
57
X-RAY DIFFRACTION
r_chiral_restr
0.104
0.2
155
X-RAY DIFFRACTION
r_gen_planes_refined
0.01
0.02
1532
X-RAY DIFFRACTION
r_gen_planes_other
0.002
0.02
315
X-RAY DIFFRACTION
r_nbd_refined
0.249
0.2
256
X-RAY DIFFRACTION
r_symmetry_nbd_other
0.205
0.2
910
X-RAY DIFFRACTION
r_nbtor_refined
0.181
0.2
539
X-RAY DIFFRACTION
r_symmetry_nbtor_other
0.091
0.2
551
X-RAY DIFFRACTION
r_xyhbond_nbd_refined
0.199
0.2
80
X-RAY DIFFRACTION
r_metal_ion_refined
0.085
0.2
3
X-RAY DIFFRACTION
r_symmetry_nbd_refined
0.25
0.2
12
X-RAY DIFFRACTION
r_nbd_other
0.163
0.2
75
X-RAY DIFFRACTION
r_symmetry_xyhbond_nbd_refined
0.267
0.2
30
X-RAY DIFFRACTION
r_mcbond_it
2.188
2.222
555
X-RAY DIFFRACTION
r_mcbond_other
2.185
2.221
555
X-RAY DIFFRACTION
r_mcangle_it
3.091
4.003
706
X-RAY DIFFRACTION
r_mcangle_other
3.092
4.007
707
X-RAY DIFFRACTION
r_scbond_it
3.167
2.633
611
X-RAY DIFFRACTION
r_scbond_other
3.12
2.589
562
X-RAY DIFFRACTION
r_scangle_it
4.784
4.702
891
X-RAY DIFFRACTION
r_scangle_other
4.771
4.594
817
X-RAY DIFFRACTION
r_lrange_it
6.611
25.638
1397
X-RAY DIFFRACTION
r_lrange_other
6.518
23.41
1329
LS refinement shell
Refine-ID: X-RAY DIFFRACTION / Total num. of bins used: 20
Resolution (Å)
Rfactor Rfree
Num. reflection Rfree
Rfactor Rwork
Num. reflection Rwork
Rfactor all
Num. reflection all
Fsc free
Fsc work
% reflection obs (%)
WRfactor Rwork
1.42-1.457
0.337
100
0.291
1510
0.294
1610
0.929
0.947
100
0.25
1.457-1.497
0.349
83
0.272
1490
0.276
1573
0.925
0.956
100
0.226
1.497-1.54
0.296
72
0.252
1437
0.254
1509
0.948
0.96
100
0.211
1.54-1.587
0.265
80
0.243
1396
0.244
1476
0.957
0.962
100
0.201
1.587-1.639
0.246
76
0.223
1356
0.224
1433
0.964
0.969
99.9302
0.186
1.639-1.697
0.262
70
0.234
1327
0.235
1398
0.958
0.966
99.9285
0.196
1.697-1.761
0.324
65
0.223
1283
0.228
1349
0.932
0.967
99.9259
0.192
1.761-1.832
0.274
56
0.212
1234
0.215
1291
0.952
0.971
99.9225
0.187
1.832-1.914
0.236
63
0.22
1188
0.22
1251
0.968
0.97
100
0.203
1.914-2.007
0.247
64
0.184
1133
0.187
1198
0.965
0.978
99.9165
0.178
2.007-2.115
0.196
55
0.178
1075
0.179
1130
0.975
0.98
100
0.178
2.115-2.243
0.219
57
0.177
1029
0.179
1087
0.971
0.981
99.908
0.184
2.243-2.397
0.146
58
0.169
969
0.168
1028
0.983
0.983
99.9027
0.184
2.397-2.589
0.204
68
0.169
880
0.171
949
0.975
0.982
99.8946
0.185
2.589-2.834
0.21
40
0.171
856
0.172
896
0.97
0.982
100
0.197
2.834-3.167
0.257
53
0.168
750
0.174
805
0.962
0.98
99.7515
0.193
3.167-3.653
0.232
41
0.163
681
0.166
724
0.96
0.985
99.7238
0.199
3.653-4.464
0.177
35
0.152
588
0.154
625
0.98
0.985
99.68
0.198
4.464-6.272
0.202
18
0.224
476
0.223
501
0.974
0.974
98.6028
0.292
6.272-38.945
0.299
13
0.305
299
0.305
313
0.968
0.939
99.6805
0.399
+
About Yorodumi
-
News
-
Feb 9, 2022. New format data for meta-information of EMDB entries
New format data for meta-information of EMDB entries
Version 3 of the EMDB header file is now the official format.
The previous official version 1.9 will be removed from the archive.
In the structure databanks used in Yorodumi, some data are registered as the other names, "COVID-19 virus" and "2019-nCoV". Here are the details of the virus and the list of structure data.
Jan 31, 2019. EMDB accession codes are about to change! (news from PDBe EMDB page)
EMDB accession codes are about to change! (news from PDBe EMDB page)
The allocation of 4 digits for EMDB accession codes will soon come to an end. Whilst these codes will remain in use, new EMDB accession codes will include an additional digit and will expand incrementally as the available range of codes is exhausted. The current 4-digit format prefixed with “EMD-” (i.e. EMD-XXXX) will advance to a 5-digit format (i.e. EMD-XXXXX), and so on. It is currently estimated that the 4-digit codes will be depleted around Spring 2019, at which point the 5-digit format will come into force.
The EM Navigator/Yorodumi systems omit the EMD- prefix.
Related info.:Q: What is EMD? / ID/Accession-code notation in Yorodumi/EM Navigator
Yorodumi is a browser for structure data from EMDB, PDB, SASBDB, etc.
This page is also the successor to EM Navigator detail page, and also detail information page/front-end page for Omokage search.
The word "yorodu" (or yorozu) is an old Japanese word meaning "ten thousand". "mi" (miru) is to see.
Related info.:EMDB / PDB / SASBDB / Comparison of 3 databanks / Yorodumi Search / Aug 31, 2016. New EM Navigator & Yorodumi / Yorodumi Papers / Jmol/JSmol / Function and homology information / Changes in new EM Navigator and Yorodumi
 Movie
 Controller


 Yorodumi
 Open data

 Basic information
 Components
 Keywords
 Function and homology information

 Gallus gallus (chicken)
 X-RAY DIFFRACTION /
 Authors
 Citation
 Structure visualization
 Downloads & links
 Other downloads


 PDBj
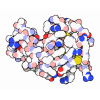
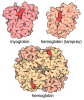
 Assembly

 Mass: 22.990 Da / Num. of mol.: 1 / Source method: obtained synthetically / Formula: Na
 Mass: 62.005 Da / Num. of mol.: 3 / Source method: obtained synthetically / Formula: NO3
 Mass: 18.015 Da / Num. of mol.: 126 / Source method: isolated from a natural source / Formula: H2O
 Sample preparation
 / Beamline: 11.2C / Wavelength: 1 Å
 Processing