Movie
 Controller


+ Open data

- Basic information
| Entry | Database: PDB / ID: 12cx | ||||||||||||
|---|---|---|---|---|---|---|---|---|---|---|---|---|---|
| Title | D189K thrombin inhibited with D-Phe-Pro-Phe-Chloromethylketone | ||||||||||||
 Components |
| ||||||||||||
 Keywords | BLOOD CLOTTING / serine protease / inhibitor | ||||||||||||
| Function / homology |  Function and homology information negative regulation of astrocyte differentiation / thrombospondin receptor activity / thrombin / thrombin-activated receptor signaling pathway / Defective factor XII causes hereditary angioedema / regulation of blood coagulation / neutrophil-mediated killing of gram-negative bacterium / positive regulation of phospholipase C-activating G protein-coupled receptor signaling pathway / Platelet Aggregation (Plug Formation) / Defective F8 cleavage by thrombin ...negative regulation of astrocyte differentiation / thrombospondin receptor activity / thrombin / thrombin-activated receptor signaling pathway / Defective factor XII causes hereditary angioedema / regulation of blood coagulation / neutrophil-mediated killing of gram-negative bacterium / positive regulation of phospholipase C-activating G protein-coupled receptor signaling pathway / Platelet Aggregation (Plug Formation) / Defective F8 cleavage by thrombin / positive regulation of collagen biosynthetic process / Fibrin formation / negative regulation of platelet activation / negative regulation of blood coagulation / negative regulation of fibrinolysis / positive regulation of blood coagulation / negative regulation of proteolysis / Transport of gamma-carboxylated protein precursors from the endoplasmic reticulum to the Golgi apparatus / fibrinolysis / Gamma-carboxylation of protein precursors / Amplification and propagation of coagulation cascade / Removal of aminoterminal propeptides from gamma-carboxylated proteins / Initiation of coagulation cascade / regulation of cytosolic calcium ion concentration / negative regulation of cytokine production involved in inflammatory response / Regulation of clotting cascade / Regulation of Complement cascade / positive regulation of receptor signaling pathway via JAK-STAT / positive regulation of release of sequestered calcium ion into cytosol / Cell surface interactions at the vascular wall / growth factor activity / Peptide ligand-binding receptors / response to wounding / lipopolysaccharide binding / platelet activation / positive regulation of protein localization to nucleus / Golgi lumen / blood coagulation / Regulation of Insulin-like Growth Factor (IGF) transport and uptake by Insulin-like Growth Factor Binding Proteins (IGFBPs) / positive regulation of reactive oxygen species metabolic process / heparin binding / Thrombin signalling through proteinase activated receptors (PARs) / Dengue Virus-Host Interactions / antimicrobial humoral immune response mediated by antimicrobial peptide / blood microparticle / extracellular matrix / G alpha (q) signalling events / cell surface receptor signaling pathway / receptor ligand activity / endoplasmic reticulum lumen / serine-type endopeptidase activity / signaling receptor binding / calcium ion binding / proteolysis / extracellular exosome / extracellular region / plasma membrane Similarity search - Function | ||||||||||||
| Biological species |  Homo sapiens (human) | ||||||||||||
| Method |  X-RAY DIFFRACTION / SYNCHROTRON / MOLECULAR REPLACEMENT / Resolution: 1.42 Å | ||||||||||||
 Authors | Friet, T. / Mohammed, B.M. / Sukumar, N. / Di Cera, E. | ||||||||||||
| Funding support |  United States, 3items
| ||||||||||||
 Citation | Journal: J.Thromb.Haemost. / Year: 2026 Title: Structural analysis of the primary specificity of thrombin. Authors: Friet, T. / Mikhail, G. / Mohammed, B.M. / Pelc, L.A. / Dei Rossi, A. / Korolev, S. / Di Cera, E. | ||||||||||||
| History |
|
- Structure visualization
| Structure viewer | Molecule: MolmilJmol/JSmol |
|---|
- Downloads & links
-Download
| PDBx/mmCIF format | 12cx.cif.gz | 94.4 KB | Display | PDBx/mmCIF format |
|---|---|---|---|---|
| PDB format | pdb12cx.ent.gz | Display | PDB format | |
| PDBx/mmJSON format | 12cx.json.gz | Tree view | PDBx/mmJSON format | |
| Others |  Other downloads |
-Validation report
| Arichive directory | https://data.pdbj.org/pub/pdb/validation_reports/2c/12cxftp://data.pdbj.org/pub/pdb/validation_reports/2c/12cx | HTTPS FTP |
|---|
-Related structure data
| Related structure data |  11riC  11rjC  11rkC  11vgC  11xwC  11yhC  12cwC  12cyC  12itC C: citing same article ( |
|---|---|
| Similar structure data |
-Links
 PDBj
- Assembly
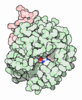
| Deposited unit | 
| ||||||||||||
|---|---|---|---|---|---|---|---|---|---|---|---|---|---|
| 1 |
| ||||||||||||
| Unit cell |
| ||||||||||||
| Components on special symmetry positions |
|
-Components
| #1: Protein/peptide | Mass: 5641.175 Da / Num. of mol.: 1 Source method: isolated from a genetically manipulated source Source: (gene. exp.) Homo sapiens (human) / Gene: F2 / Production host:  Mesocricetus auratus (golden hamster) / References: UniProt: P00734 |
|---|---|
| #2: Protein | Mass: 31439.127 Da / Num. of mol.: 1 / Mutation: D519K Source method: isolated from a genetically manipulated source Source: (gene. exp.) Homo sapiens (human) / Gene: F2 / Production host: Mesocricetus auratus (golden hamster) / References: UniProt: P00734 |
| #3: Chemical | ChemComp-A1DFL / Mass: 443.966 Da / Num. of mol.: 1 / Source method: obtained synthetically / Formula: C24H30ClN3O3 / Feature type: SUBJECT OF INVESTIGATION |
| #4: Sugar | ChemComp-NAG /   Type: D-saccharide, beta linking / Mass: 221.208 Da / Num. of mol.: 1 / Source method: isolated from a natural source / Formula: C8H15NO6 |
| #5: Water | ChemComp-HOH /  Mass: 18.015 Da / Num. of mol.: 193 / Source method: isolated from a natural source / Formula: H2O |
| Has ligand of interest | Y |
| Has protein modification | Y |
-Experimental details
-Experiment
| Experiment | Method: X-RAY DIFFRACTION / Number of used crystals: 1 |
|---|
- Sample preparation
| Crystal | Density Matthews: 2.33 Å3/Da / Density % sol: 47.27 % |
|---|---|
| Crystal grow | Temperature: 293.15 K / Method: vapor diffusion, hanging drop Details: 0.2 M Sodium chloride, 0.1 M BIS-TRIS pH 5.5, 25% w/v Polyethylene glycol 3,350 |
-Data collection
| Diffraction | Mean temperature: 100 K / Serial crystal experiment: N |
|---|---|
| Diffraction source | Source: SYNCHROTRON / Site: NSLS-II / Beamline: 17-ID-2 / Wavelength: 0.97934 Å |
| Detector | Type: DECTRIS EIGER X 16M / Detector: PIXEL / Date: Apr 4, 2025 |
| Radiation | Protocol: SINGLE WAVELENGTH / Monochromatic (M) / Laue (L): M / Scattering type: x-ray |
| Radiation wavelength | Wavelength: 0.97934 Å / Relative weight: 1 |
| Reflection | Resolution: 1.42→47.63 Å / Num. obs: 62265 / % possible obs: 99.5 % / Redundancy: 6.8 % / Biso Wilson estimate: 11.25 Å2 / CC1/2: 0.986 / Rmerge(I) obs: 0.18 / Rpim(I) all: 0.074 / Rrim(I) all: 0.195 / Χ2: 0.93 / Net I/σ(I): 5.8 / Num. measured all: 423993 |
| Reflection shell | Resolution: 1.42→1.5 Å / % possible obs: 98.2 % / Redundancy: 6.3 % / Rmerge(I) obs: 0.881 / Num. measured all: 56500 / Num. unique obs: 8912 / CC1/2: 0.432 / Rpim(I) all: 0.375 / Rrim(I) all: 0.959 / Χ2: 0.83 / Net I/σ(I) obs: 2.2 |
- Processing
| Software |
| |||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||
|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|
| Refinement | Method to determine structure: MOLECULAR REPLACEMENT / Resolution: 1.42→43.41 Å / SU ML: 0.1828 / Cross valid method: FREE R-VALUE / σ(F): 1.35 / Phase error: 24.5005 Stereochemistry target values: GeoStd + Monomer Library + CDL v1.2
| |||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||
| Solvent computation | Shrinkage radii: 0.9 Å / VDW probe radii: 1.1 Å / Solvent model: FLAT BULK SOLVENT MODEL | |||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||
| Displacement parameters | Biso mean: 16.64 Å2 | |||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||
| Refinement step | Cycle: LAST / Resolution: 1.42→43.41 Å
| |||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||
| Refine LS restraints |
| |||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||
| LS refinement shell |
|