Movie
 Controller


[English] 日本語
 Yorodumi
- PDB-38we: Crystal structure of the CFTR inhibitory factor Cif with the F164... -
+ Open data

- Basic information
| Entry | Database: PDB / ID: 38we | ||||||||||||
|---|---|---|---|---|---|---|---|---|---|---|---|---|---|
| Title | Crystal structure of the CFTR inhibitory factor Cif with the F164A mutation | ||||||||||||
 Components | CFTR inhibitory factor | ||||||||||||
 Keywords | HYDROLASE / Epoxide hydrolase / Virulence factor | ||||||||||||
| Function / homology | soluble epoxide hydrolase / Alpha/beta hydrolase family / epoxide hydrolase activity / Alpha/beta hydrolase fold-1 / host cell / Alpha/Beta hydrolase fold / extracellular region / CFTR inhibitory factor Function and homology information | ||||||||||||
| Biological species |  Pseudomonas aeruginosa UCBPP-PA14 (bacteria) | ||||||||||||
| Method |  X-RAY DIFFRACTION / SYNCHROTRON / MOLECULAR REPLACEMENT / Resolution: 1.66 Å | ||||||||||||
 Authors | Nie, Z. / Madden, D.R. | ||||||||||||
| Funding support |  United States, 3items
| ||||||||||||
 Citation | Journal: To Be Published Title: Crystal structure of the CFTR inhibitory factor Cif with the F164A mutation Authors: Nie, Z. / Madden, D.R. | ||||||||||||
| History |
|
- Structure visualization
| Structure viewer | Molecule: MolmilJmol/JSmol |
|---|
- Downloads & links
-Download
| PDBx/mmCIF format | 38we.cif.gz | 285.3 KB | Display | PDBx/mmCIF format |
|---|---|---|---|---|
| PDB format | pdb38we.ent.gz | 225.6 KB | Display | PDB format |
| PDBx/mmJSON format | 38we.json.gz | Tree view | PDBx/mmJSON format | |
| Others |  Other downloads |
-Validation report
| Arichive directory | https://data.pdbj.org/pub/pdb/validation_reports/8w/38weftp://data.pdbj.org/pub/pdb/validation_reports/8w/38we | HTTPS FTP |
|---|
-Related structure data
| Similar structure data |
|---|
-Links
 PDBj
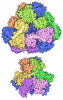
- Assembly
| Deposited unit | 
| ||||||||||||
|---|---|---|---|---|---|---|---|---|---|---|---|---|---|
| 1 | 
| ||||||||||||
| 2 | 
| ||||||||||||
| Unit cell |
| ||||||||||||
| Components on special symmetry positions |
|
-Components
| #1: Protein | Mass: 33259.734 Da / Num. of mol.: 4 / Mutation: F164A Source method: isolated from a genetically manipulated source Source: (gene. exp.) Pseudomonas aeruginosa UCBPP-PA14 (bacteria)Gene: cif, PA14_26090 / Plasmid: pET16b Details (production host): N-terminal decahistidine-SUMO fusion Production host:  Escherichia coli BL21(DE3) (bacteria) / References: UniProt: A0A0H2ZD27, soluble epoxide hydrolase #2: Chemical | ChemComp-CA /   Mass: 40.078 Da / Num. of mol.: 6 / Source method: obtained synthetically / Formula: Ca #3: Water | ChemComp-HOH / |  Mass: 18.015 Da / Num. of mol.: 1331 / Source method: isolated from a natural source / Formula: H2O Has ligand of interest | N | Has protein modification | Y | |
|---|
-Experimental details
-Experiment
| Experiment | Method: X-RAY DIFFRACTION / Number of used crystals: 1 |
|---|
- Sample preparation
| Crystal | Density Matthews: 2.34 Å3/Da / Density % sol: 47.56 % |
|---|---|
| Crystal grow | Temperature: 291 K / Method: vapor diffusion, hanging drop / pH: 5 Details: 17% (w/v) PEG 8000, 175 mM calcium chloride, 100 mM sodium acetate, pH 5.0 |
-Data collection
| Diffraction | Mean temperature: 100 K / Serial crystal experiment: N |
|---|---|
| Diffraction source | Source: SYNCHROTRON / Site: NSLS-II / Beamline: 17-ID-1 / Wavelength: 0.92019 Å |
| Detector | Type: DECTRIS EIGER X 9M / Detector: PIXEL / Date: Mar 14, 2025 / Details: KB bimorph mirrors |
| Radiation | Monochromator: Si(111) DCM / Protocol: SINGLE WAVELENGTH / Monochromatic (M) / Laue (L): M / Scattering type: x-ray |
| Radiation wavelength | Wavelength: 0.92019 Å / Relative weight: 1 |
| Reflection | Resolution: 1.66→43.72 Å / Num. obs: 144615 / % possible obs: 99.89 % / Redundancy: 4.4 % / Biso Wilson estimate: 19.35 Å2 / CC1/2: 0.998 / CC star: 0.999 / Rmerge(I) obs: 0.0911 / Rpim(I) all: 0.0483 / Rrim(I) all: 0.1035 / Net I/σ(I): 8.73 |
| Reflection shell | Resolution: 1.66→1.72 Å / Redundancy: 4.4 % / Rmerge(I) obs: 0.6669 / Mean I/σ(I) obs: 1.61 / Num. unique obs: 14442 / CC1/2: 0.685 / CC star: 0.902 / Rpim(I) all: 0.357 / Rrim(I) all: 0.7592 / % possible all: 99.93 |
- Processing
| Software |
| |||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||
|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|
| Refinement | Method to determine structure: MOLECULAR REPLACEMENT / Resolution: 1.66→43.72 Å / SU ML: 0.2357 / Cross valid method: FREE R-VALUE / σ(F): 1.33 / Phase error: 19.3263 Stereochemistry target values: GeoStd + Monomer Library + CDL v1.2
| |||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||
| Solvent computation | Shrinkage radii: 0.9 Å / VDW probe radii: 1.1 Å / Solvent model: FLAT BULK SOLVENT MODEL | |||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||
| Displacement parameters | Biso mean: 21.59 Å2 | |||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||
| Refinement step | Cycle: LAST / Resolution: 1.66→43.72 Å
| |||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||
| Refine LS restraints |
| |||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||
| LS refinement shell |
|