 Movie
 Controller


[English] 日本語
 Yorodumi
- PDB-25ud: Crystal structure of SktA in complex with 5,7-dibromo-L-tryptopha... -
+ Open data

- Basic information
| Entry | Database: PDB / ID: 25ud | ||||||
|---|---|---|---|---|---|---|---|
| Title | Crystal structure of SktA in complex with 5,7-dibromo-L-tryptophan and Fe2+ | ||||||
 Components | Iron-containing redox enzyme family protein | ||||||
 Keywords | OXIDOREDUCTASE / skatole synthase / 5 / 7-dibromo-L-tryptophan / diiron oxidases | ||||||
| Function / homology | 5-bromoskatole synthase / Haem oxygenase-like, multi-helical / Chem-67I / : / 5-bromoskatole synthase Function and homology information | ||||||
| Biological species |  Nostoc punctiforme NIES-2108 (bacteria) | ||||||
| Method |  X-RAY DIFFRACTION / SYNCHROTRON / MOLECULAR REPLACEMENT / Resolution: 2.5 Å | ||||||
 Authors | Li, H. / Liu, J. / Huang, J.-W. / Chen, C.-C. / Guo, R.-T. | ||||||
| Funding support |  China, 1items
| ||||||
 Citation | Journal: Acs Catalysis / Year: 2026 Title: Structural Insights and Functional Expansion of an Aerobic Skatole Synthase Authors: Li, H. / Huang, J.W. / Liu, J. / Zhang, N. / Dai, L. / Li, S. / Chen, C.C. / Guo, R.T. | ||||||
| History |
|
- Structure visualization
| Structure viewer | Molecule: MolmilJmol/JSmol |
|---|
- Downloads & links
-Download
| PDBx/mmCIF format | 25ud.cif.gz | 197.6 KB | Display | PDBx/mmCIF format |
|---|---|---|---|---|
| PDB format | pdb25ud.ent.gz | 157.6 KB | Display | PDB format |
| PDBx/mmJSON format | 25ud.json.gz | Tree view | PDBx/mmJSON format | |
| Others |  Other downloads |
-Validation report
| Arichive directory | https://data.pdbj.org/pub/pdb/validation_reports/5u/25udftp://data.pdbj.org/pub/pdb/validation_reports/5u/25ud | HTTPS FTP |
|---|
-Related structure data
| Related structure data |  25tzC  25uaC  25ubC  25ucC  25ueC  25ufC C: citing same article ( |
|---|---|
| Similar structure data |
-Links
 PDBj
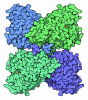
- Assembly
| Deposited unit | 
| ||||||||
|---|---|---|---|---|---|---|---|---|---|
| 1 | 
| ||||||||
| 2 | 
| ||||||||
| Unit cell |
|
-Components
| #1: Protein | Mass: 28628.359 Da / Num. of mol.: 4 Source method: isolated from a genetically manipulated source Source: (gene. exp.) Nostoc punctiforme NIES-2108 (bacteria)Gene: A6769_19210 / Production host:  Escherichia coli BL21(DE3) (bacteria) / References: UniProt: A0A367RF96 #2: Chemical | ChemComp-FE2 /   Mass: 55.845 Da / Num. of mol.: 4 / Source method: obtained synthetically / Formula: Fe / Feature type: SUBJECT OF INVESTIGATION #3: Chemical |   Mass: 362.017 Da / Num. of mol.: 3 / Source method: obtained synthetically / Formula: C11H10Br2N2O2 / Feature type: SUBJECT OF INVESTIGATION #4: Water | ChemComp-HOH / |  Mass: 18.015 Da / Num. of mol.: 141 / Source method: isolated from a natural source / Formula: H2O Has ligand of interest | Y | Has protein modification | N | |
|---|
-Experimental details
-Experiment
| Experiment | Method: X-RAY DIFFRACTION / Number of used crystals: 1 |
|---|
- Sample preparation
| Crystal | Density Matthews: 2.05 Å3/Da / Density % sol: 39.98 % |
|---|---|
| Crystal grow | Temperature: 293 K / Method: vapor diffusion, sitting drop Details: 30% PEG 4000, 0.2 M ammonium acetate, 0.1 M sodium citrate, pH 5.6 |
-Data collection
| Diffraction | Mean temperature: 100 K / Serial crystal experiment: N | ||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||
|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|
| Diffraction source | Source: SYNCHROTRON / Site: SSRF / Beamline: BL02U1 / Wavelength: 0.984 Å | ||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||
| Detector | Type: DECTRIS EIGER2 S 9M / Detector: PIXEL / Date: Jan 18, 2026 | ||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||
| Radiation | Protocol: SINGLE WAVELENGTH / Monochromatic (M) / Laue (L): M / Scattering type: x-ray | ||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||
| Radiation wavelength | Wavelength: 0.984 Å / Relative weight: 1 | ||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||
| Reflection | Resolution: 2.5→50 Å / Num. obs: 29888 / % possible obs: 96.8 % / Redundancy: 3.2 % / CC1/2: 0.991 / CC star: 0.998 / Rmerge(I) obs: 0.063 / Rpim(I) all: 0.043 / Rrim(I) all: 0.077 / Χ2: 1.451 / Net I/σ(I): 21.7 / Num. measured all: 95246 | ||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||
| Reflection shell | Diffraction-ID: 1
|
- Processing
| Software |
| ||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||
|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|
| Refinement | Method to determine structure: MOLECULAR REPLACEMENT / Resolution: 2.5→45.25 Å / Cor.coef. Fo:Fc: 0.933 / Cor.coef. Fo:Fc free: 0.891 / SU B: 11.013 / SU ML: 0.246 / Cross valid method: THROUGHOUT / ESU R Free: 0.35 / Stereochemistry target values: MAXIMUM LIKELIHOOD / Details: HYDROGENS HAVE BEEN ADDED IN THE RIDING POSITIONS
| ||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||
| Solvent computation | Ion probe radii: 0.8 Å / Shrinkage radii: 0.8 Å / VDW probe radii: 1.2 Å / Solvent model: MASK | ||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||
| Displacement parameters | Biso mean: 49.317 Å2
| ||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||
| Refinement step | Cycle: 1 / Resolution: 2.5→45.25 Å
| ||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||
| Refine LS restraints |
|